varyingPrior

Effects of varying the MAP prior

Analysing a large-ish dataset (157 non-spiked vs. 20 spiked)

Load data and prepare for simulations

Full model with default weak prior normal(0,5)

Show code
# spiked samples
ss = readLines("ecoli_sims/test/sampled.txt")

# metadata 
meta_data = data.frame(run_accession = colnames(d_p95),spiked = as.factor(sapply(colnames(d_p95), function(x) x%in% ss)))

d_p95_full = modify_metadata(se = d_p95, meta_data = meta_data)

d_p95_full <- filter_by_presence(d_p95_full, min_nonzero = 20)
Retained 487 rows after filtering
Show code
save_path = "output_rds/prior_zib95_weak.rds"
if(file.exists(save_path)){
  full_zib_95_w <- readRDS(save_path)
} else {
  system.time({
    full_zib_95_w <- glmZiBFit(d_p95_full, design, nthreads = 10)
  })
  saveRDS(full_zib_95_w, save_path)
}

top_hits(full_zib_95_w)
Found 2 tophits for spikedTRUE at alpha = 0.05 using holm 
# A tibble: 2 × 10
  Contig_name Genome_file coefficient std_error  p_value p_adjust zi_coefficient
  <chr>       <chr>             <dbl>     <dbl>    <dbl>    <dbl>          <dbl>
1 NZ_AEZI020… GCF_000194…       0.441    0.0522 3.21e-17 1.56e-14         -7.69 
2 JAHZEE0100… GCA_019421…      -0.547    0.138  7.85e- 5 3.81e- 2         -0.362
# ℹ 3 more variables: zi_std_error <dbl>, zi_p_value <dbl>, zi_p_adjust <dbl>
Show code
# true/false positive
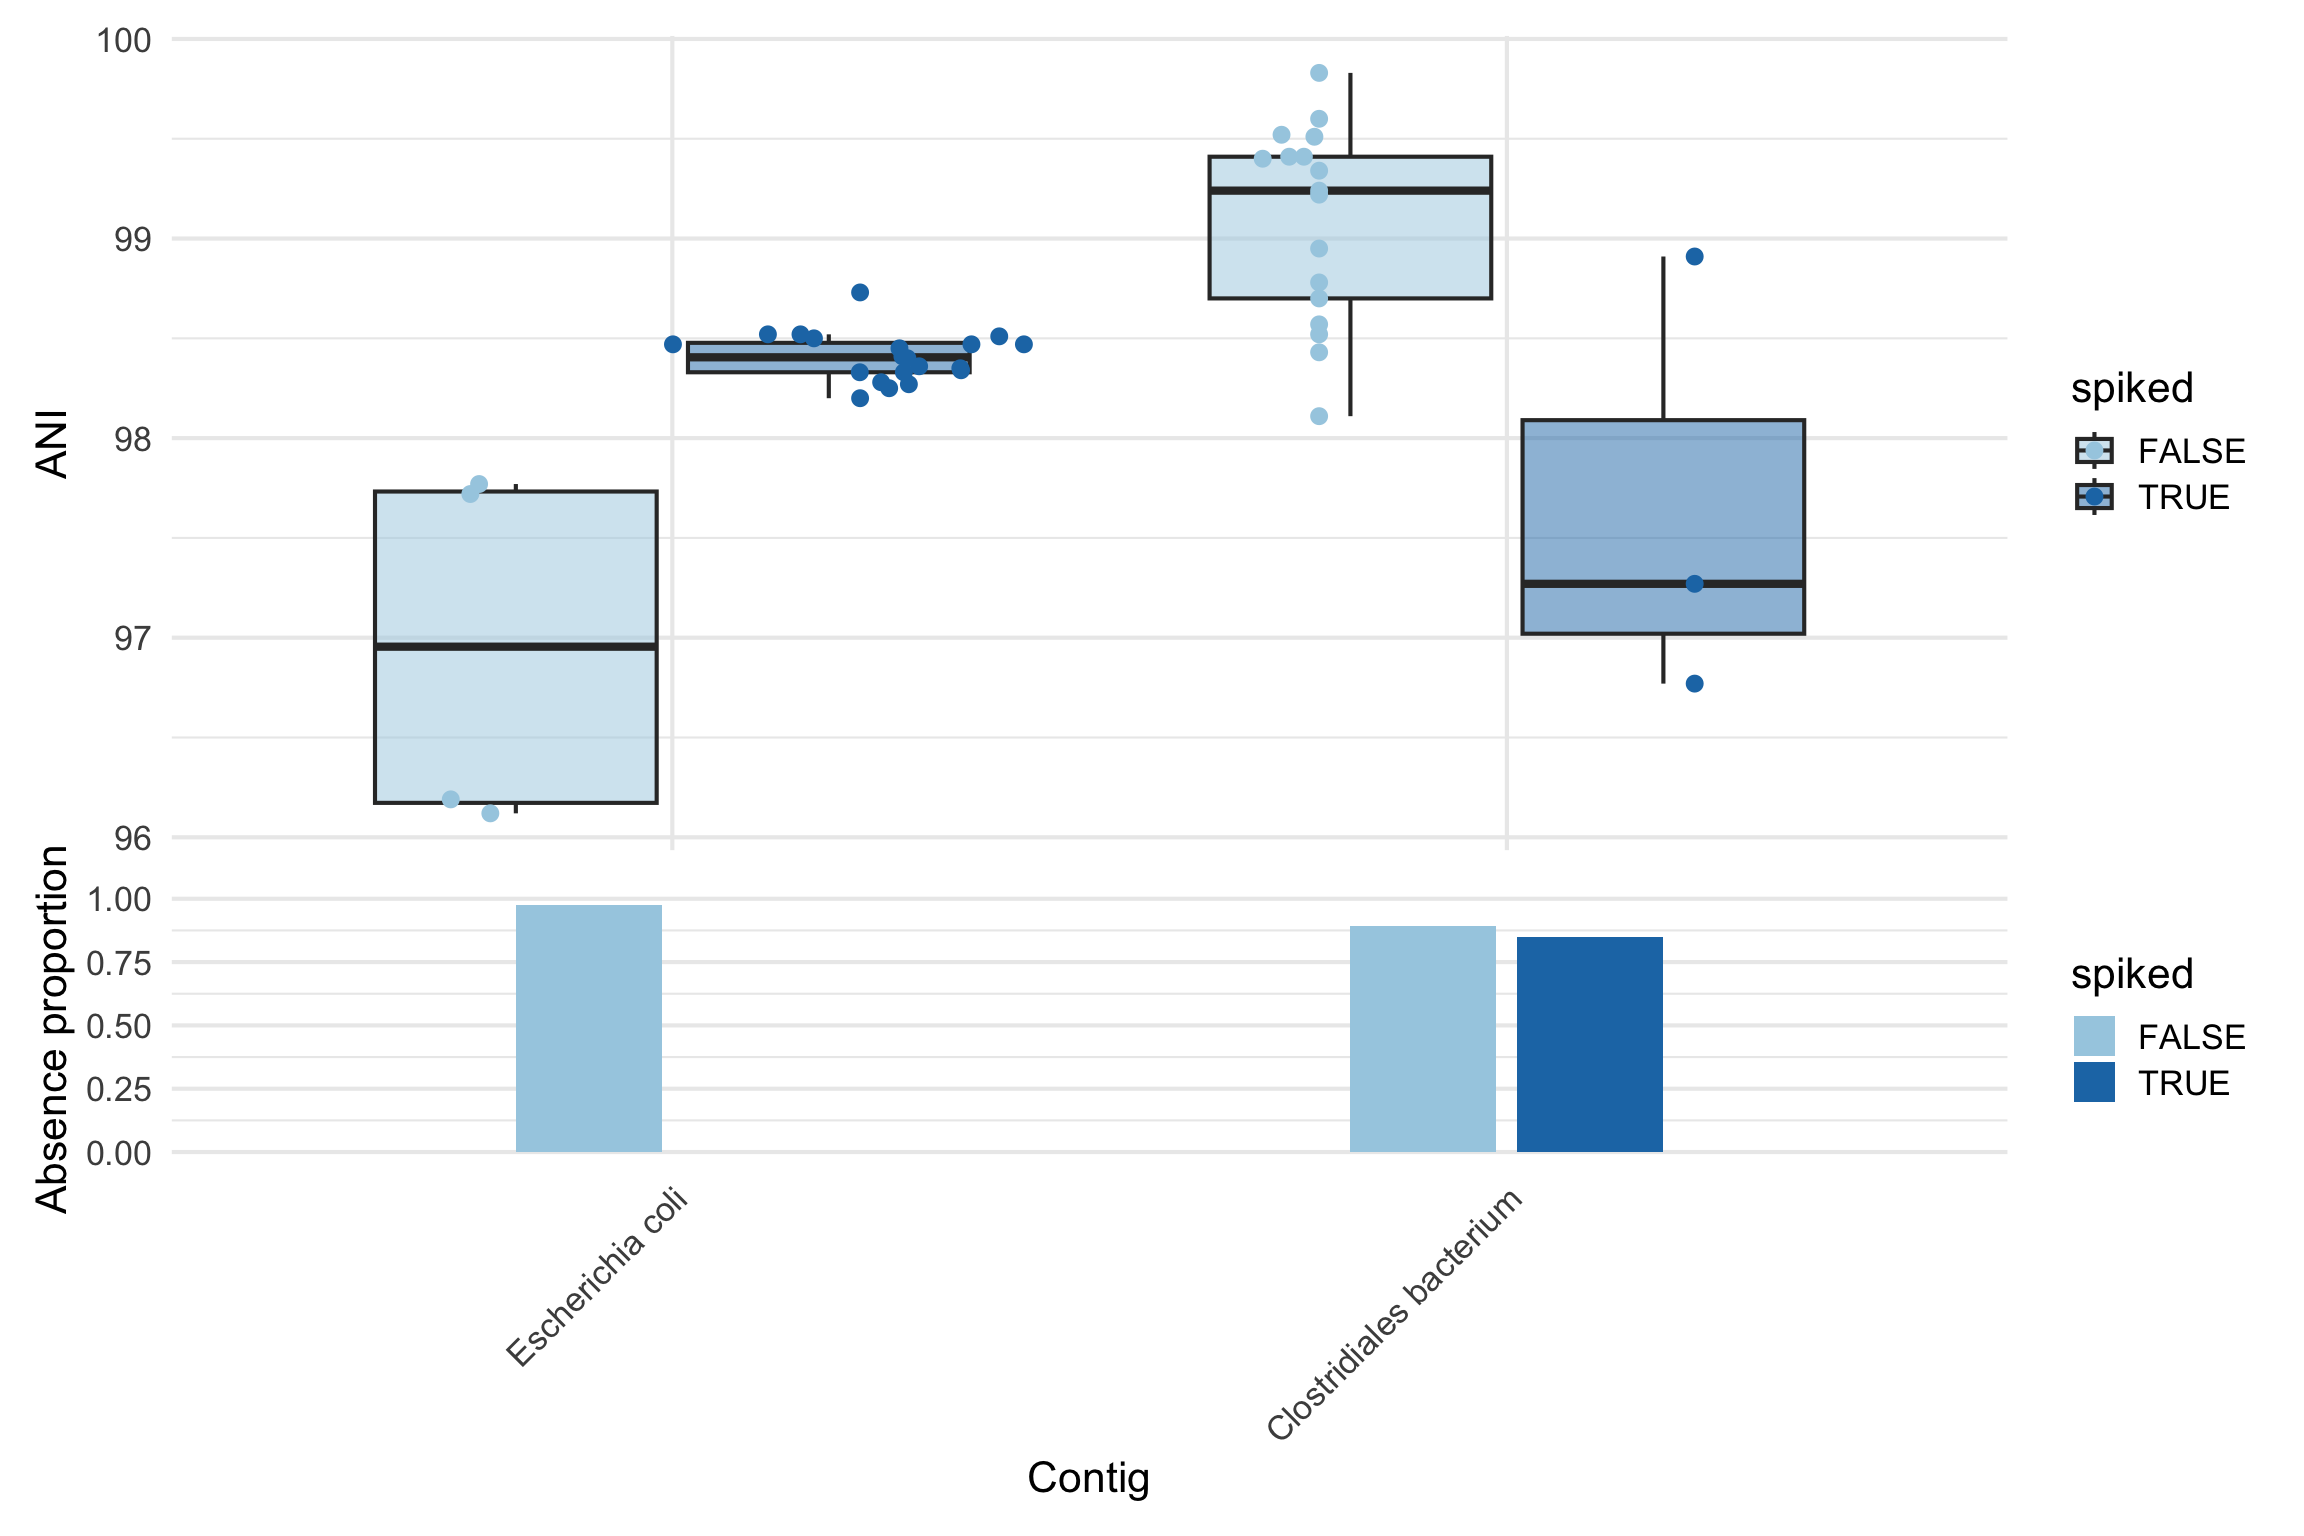
plot_ani_dist(d_p95_full, phenotype = 'spiked', contigs = top_hits(full_zib_95_w, alpha = 0.05)$Contig_name, show_points = T, plot_type = 'box')
Found 2 tophits for spikedTRUE at alpha = 0.05 using holm 
Warning: Removed 310 rows containing non-finite outside the scale range
(`stat_boxplot()`).

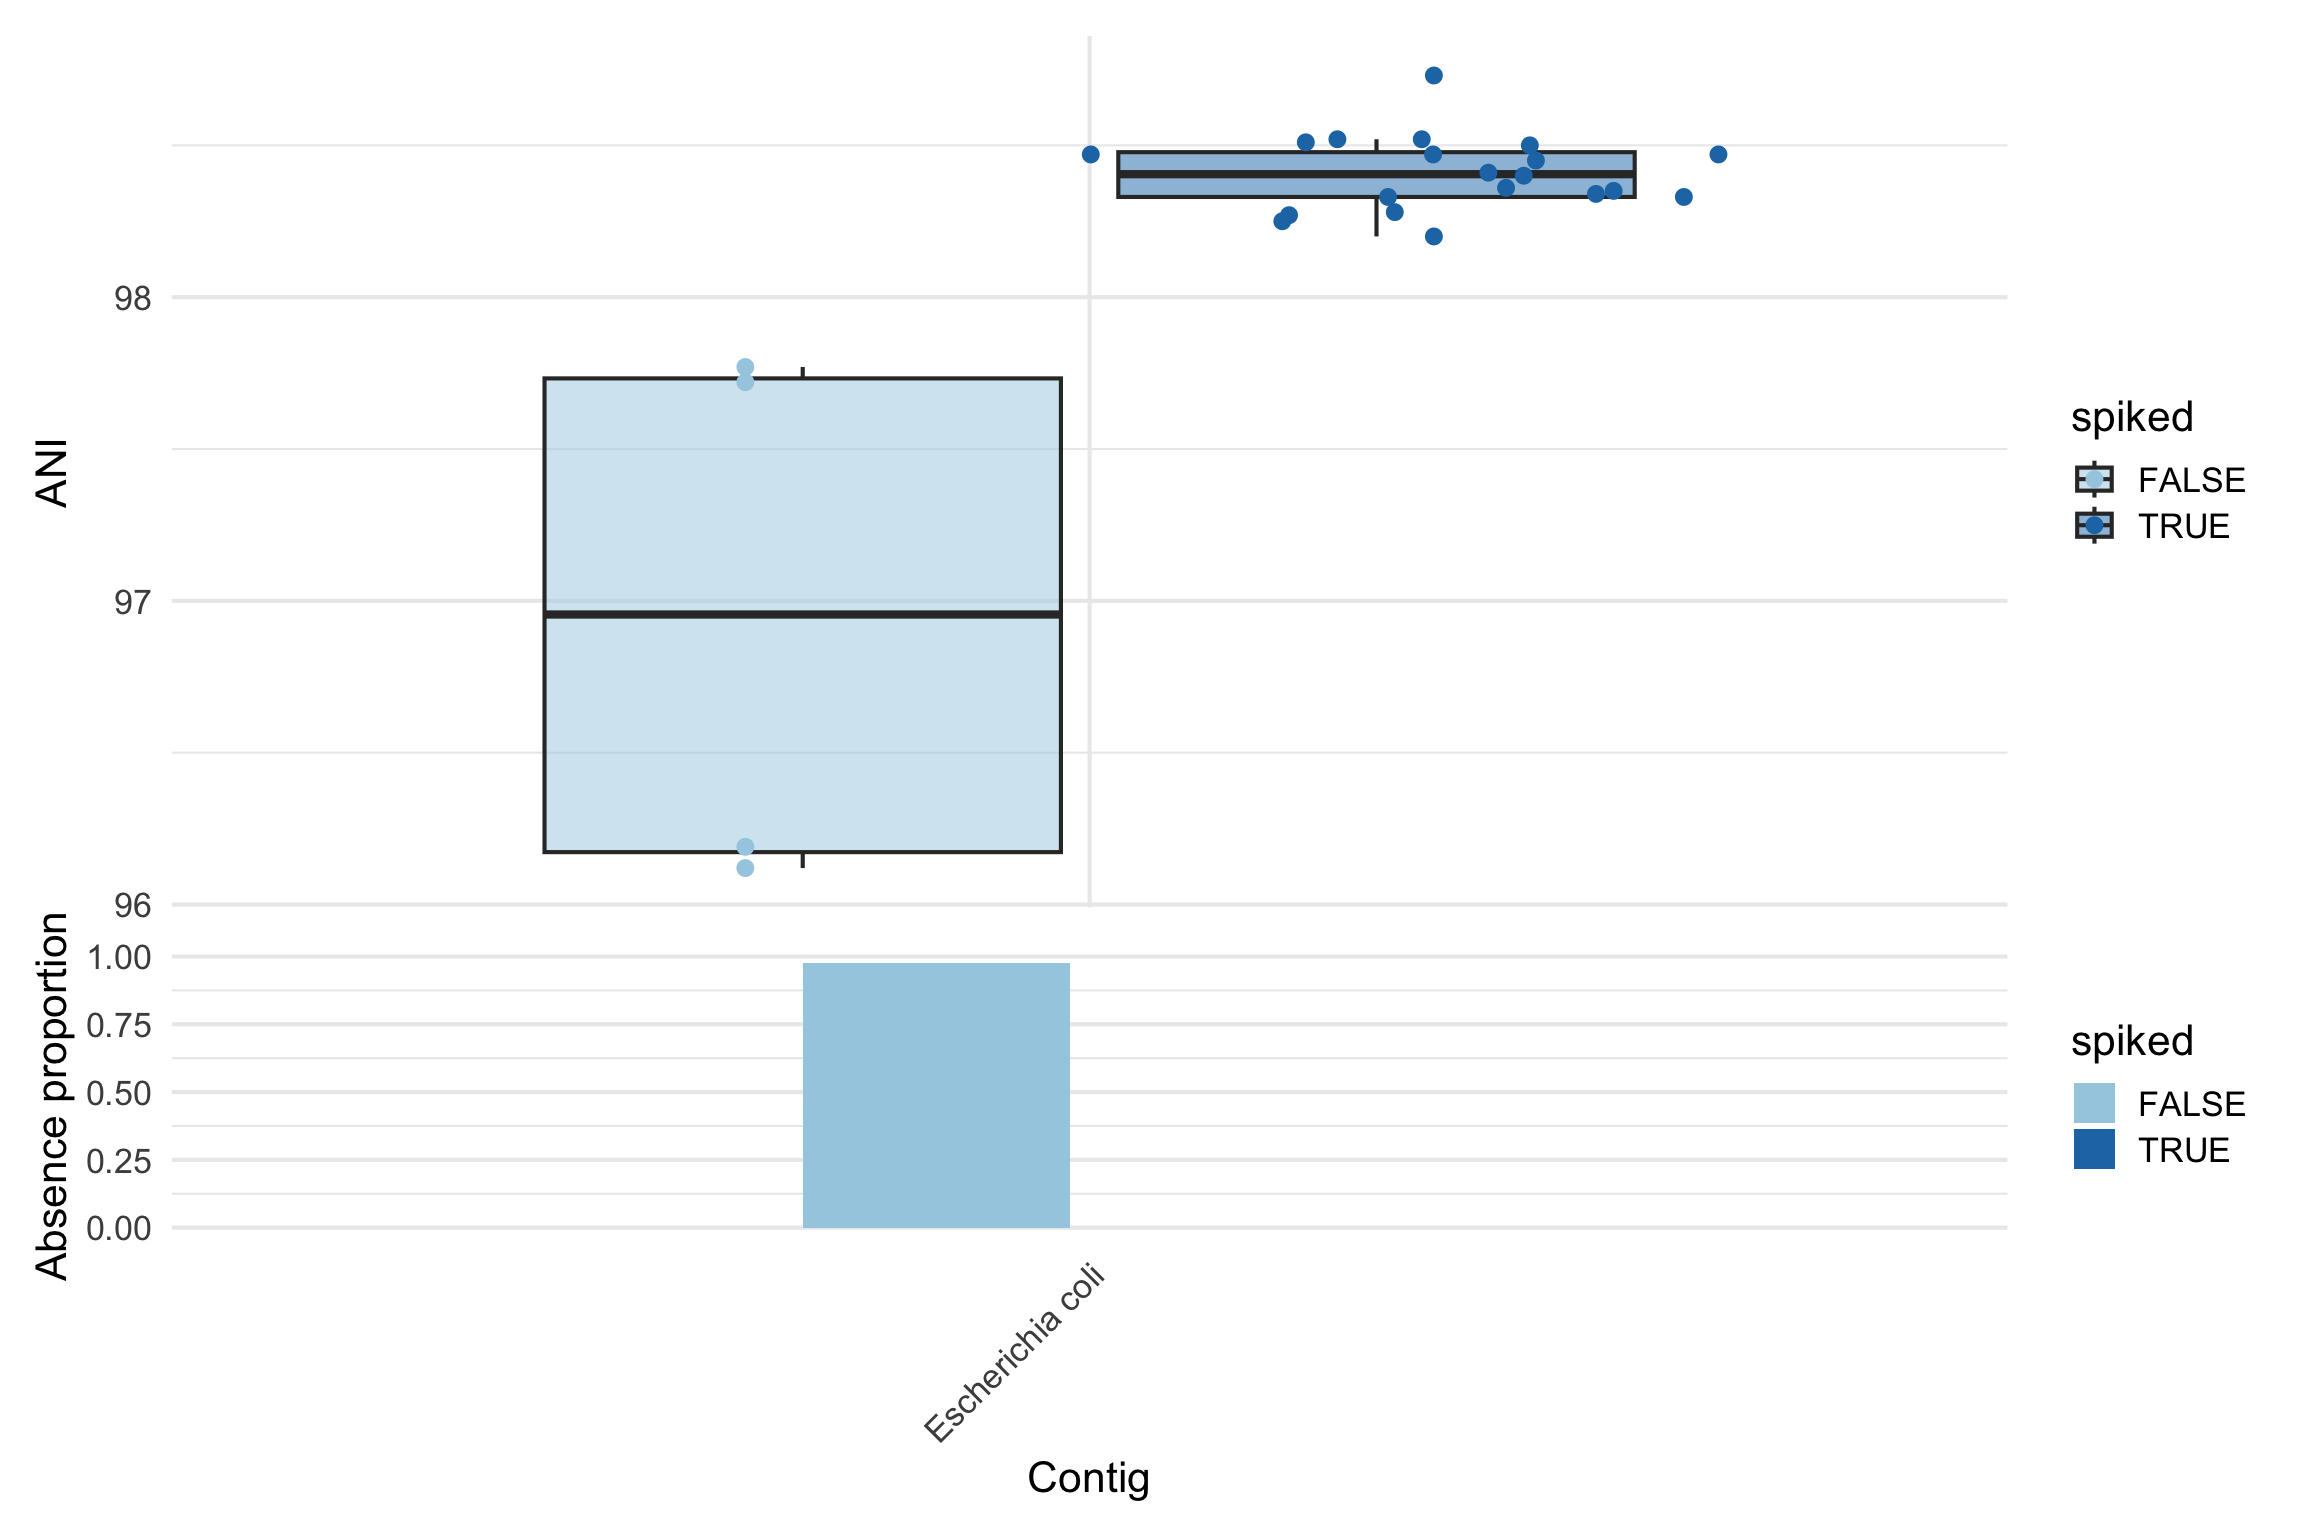
Here, Clostridiales bacterium is a false positive. Based on the box plot, it looks like an edge case with just 3 samples driving the signal.

Full model with strong prior normal(0,1)

Show code
save_path = "output_rds/prior_zib95_strong.rds"
if(file.exists(save_path)){
  full_zib_95_s <- readRDS(save_path)
} else {
  system.time({
    full_zib_95_s <- glmZiBFit(d_p95_full, design, nthreads = 10, MAP_prior = "preset_strong")
  })
  saveRDS(full_zib_95_s, save_path)
}


top_hits(full_zib_95_s)
Found 1 tophits for spikedTRUE at alpha = 0.05 using holm 
# A tibble: 1 × 10
  Contig_name Genome_file coefficient std_error  p_value p_adjust zi_coefficient
  <chr>       <chr>             <dbl>     <dbl>    <dbl>    <dbl>          <dbl>
1 NZ_AEZI020… GCF_000194…       0.446    0.0520 9.39e-18 4.57e-15          -4.01
# ℹ 3 more variables: zi_std_error <dbl>, zi_p_value <dbl>, zi_p_adjust <dbl>
Show code
# true/false positive
plot_ani_dist(d_p95_full, phenotype = 'spiked', contigs = top_hits(full_zib_95_s, alpha = 0.05)$Contig_name, show_points = T, plot_type = 'box')
Found 1 tophits for spikedTRUE at alpha = 0.05 using holm 
Warning: Removed 153 rows containing non-finite outside the scale range
(`stat_boxplot()`).

This fixes the FP issue.

Full model with ebayes prior

Show code
save_path = "output_rds/prior_zib95_ebp.rds"

ebp_full = compute_eb_priors(d_p95_full, design = design, nthreads = parallel::detectCores(), low_cutoff = 0, high_cutoff = Inf, progress = FALSE)
ebp_full
strainspy_priors object
Method:  empirical 

Priors:
 - fixef : 1 coefficients
 - fixef_zi : 1 coefficients

Example priors:
           prior    class       coef
1 normal(0,0.14)    fixef spikedTRUE
2 normal(0,2.19) fixef_zi spikedTRUE

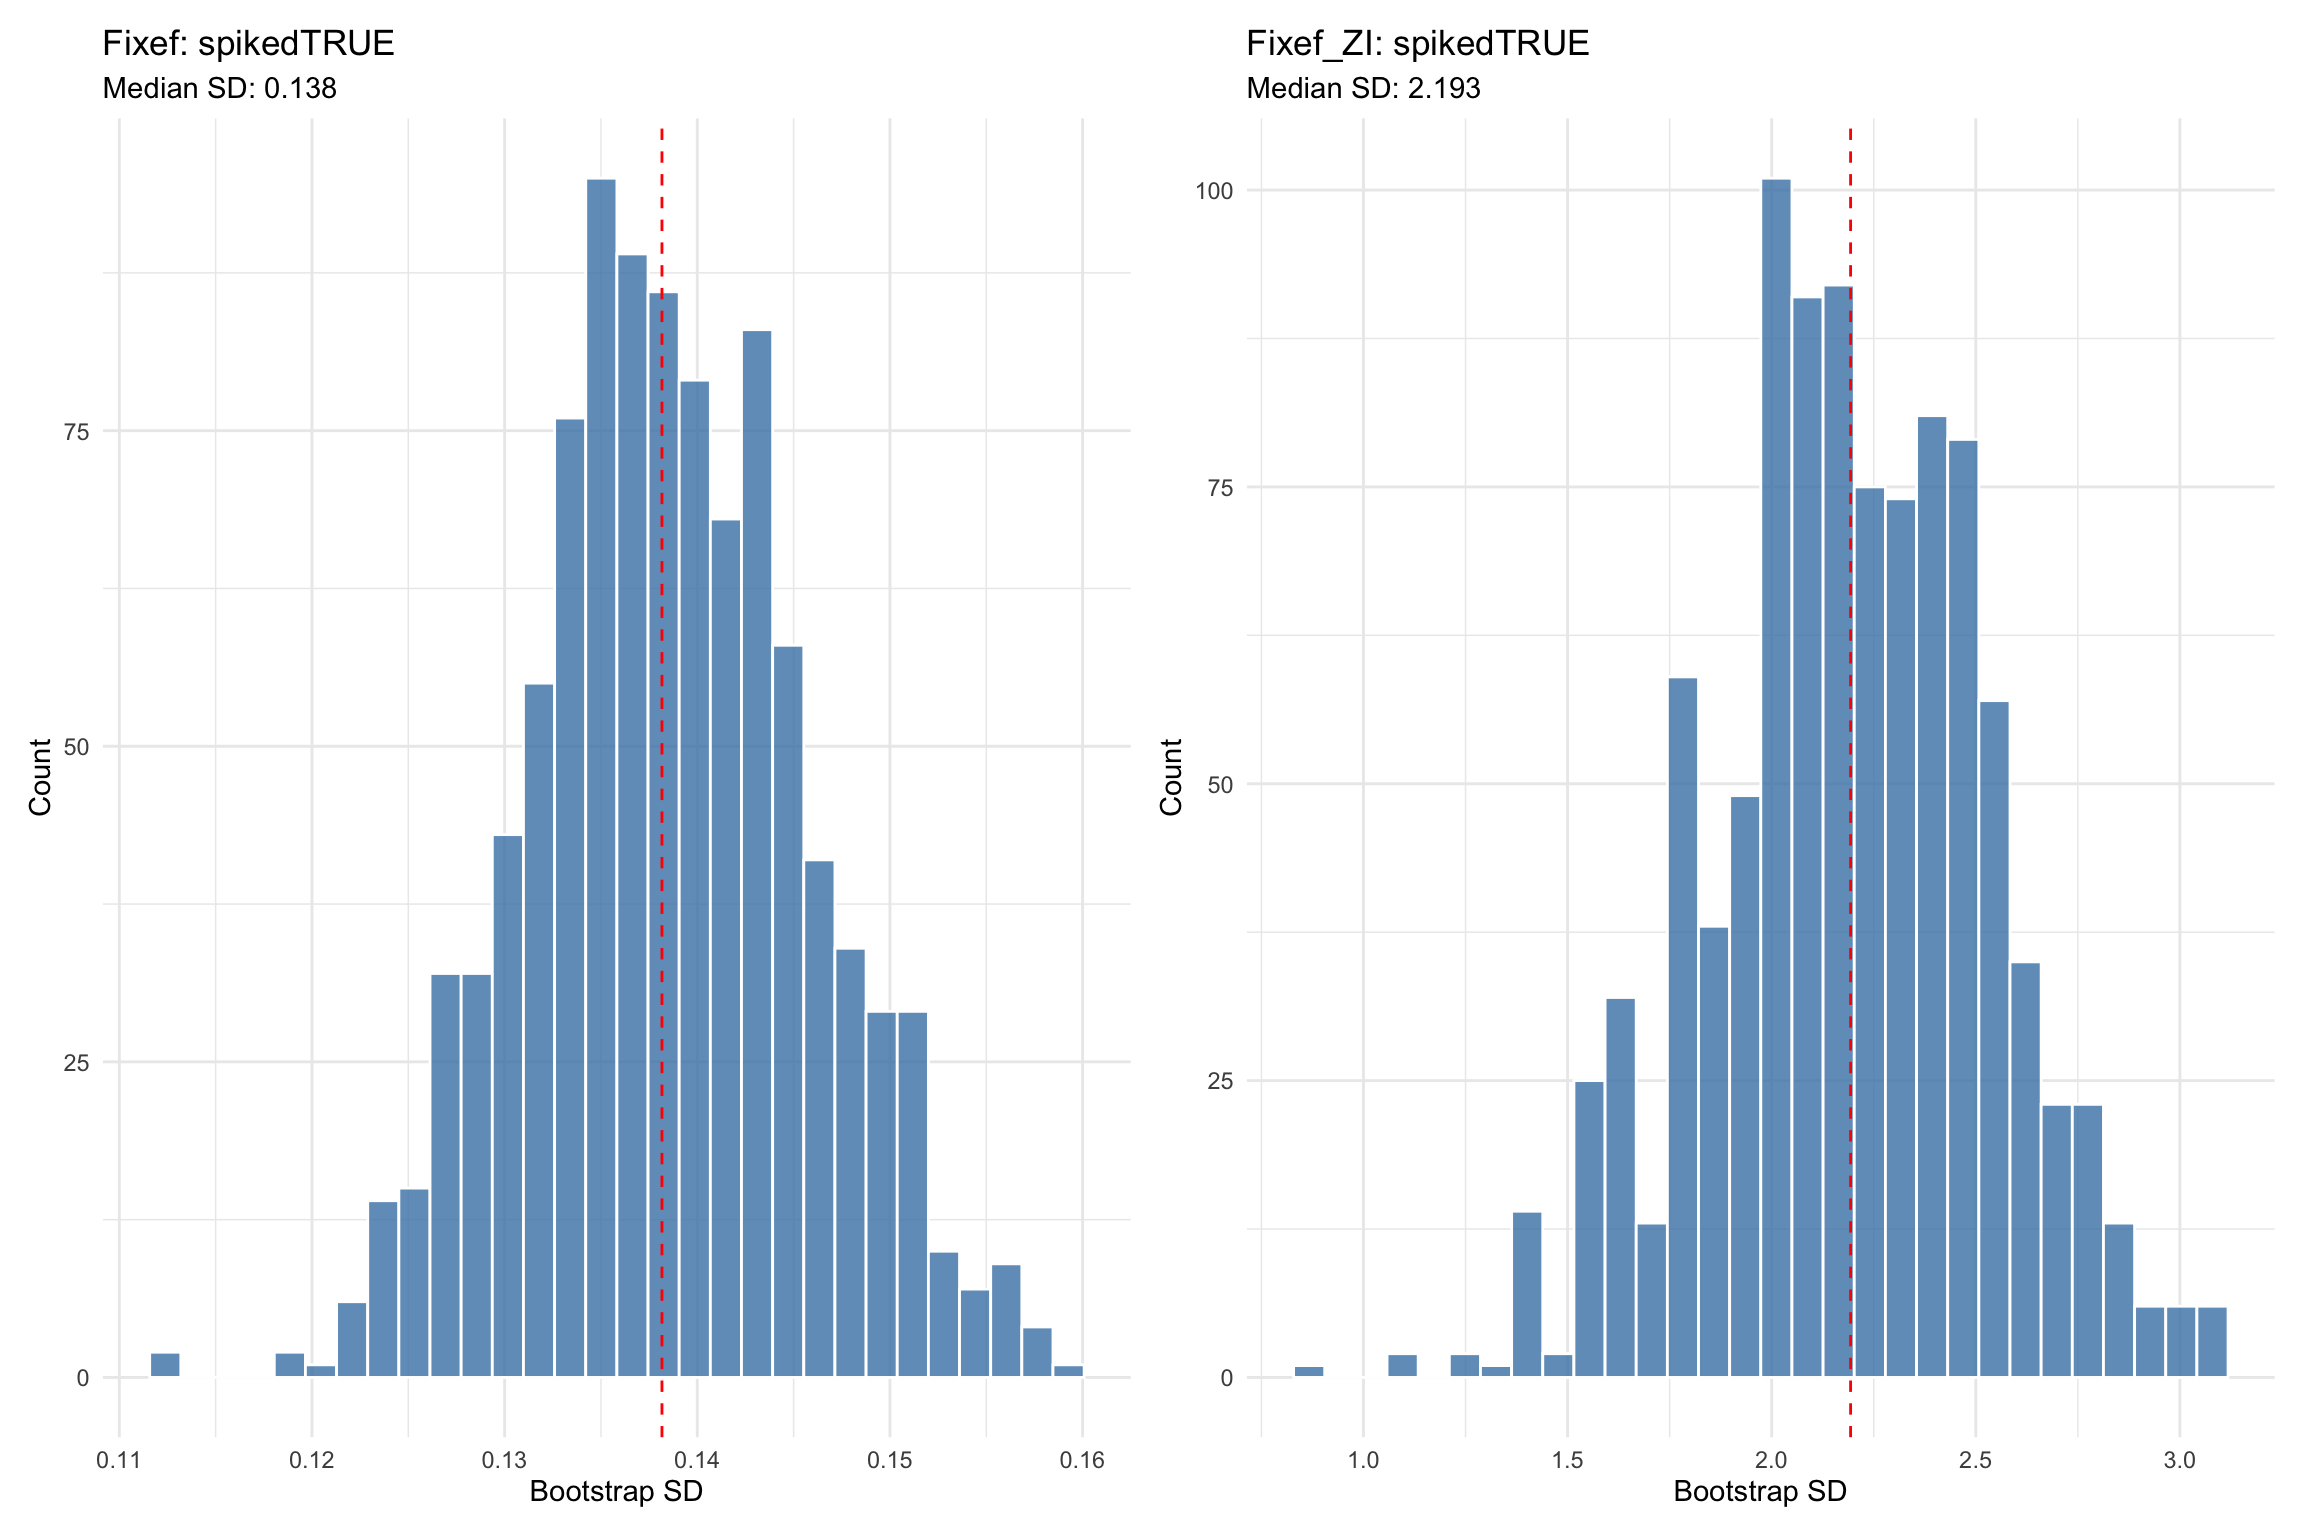
Bootstrap summary for fixef priors:
   Min. 1st Qu.  Median    Mean 3rd Qu.    Max. 
 0.1124  0.1382  0.1826  0.1837  0.2289  0.2493 

Bootstrap summary for fixef_zi priors:
   Min. 1st Qu.  Median    Mean 3rd Qu.    Max. 
 0.8669  1.1173  1.2467  1.6525  2.1932  3.0798 

Warning: 2 fixef prior(s) may be too strong (SD < 1), and 0 may be too weak (SD > 5).
Show code
plot_prior_bootstrap(object = ebp_full, "spikedTRUE")

Show code
if(file.exists(save_path)){
  full_zib_95_eb <- readRDS(save_path)
} else {
  system.time({
    full_zib_95_eb <- glmZiBFit(d_p95_full, design, nthreads = 10, MAP_prior = ebp_full)
  })
  saveRDS(full_zib_95_eb, save_path)
}

top_hits(full_zib_95_eb)
Found 1 tophits for spikedTRUE at alpha = 0.05 using holm 
# A tibble: 1 × 10
  Contig_name  Genome_file coefficient std_error p_value p_adjust zi_coefficient
  <chr>        <chr>             <dbl>     <dbl>   <dbl>    <dbl>          <dbl>
1 NZ_AEZI0200… GCF_000194…       0.197     0.135   0.145        1          -5.72
# ℹ 3 more variables: zi_std_error <dbl>, zi_p_value <dbl>, zi_p_adjust <dbl>
Show code
# This plot is redundant - same as (0,1) prior
plot_ani_dist(d_p95_full, phenotype = 'spiked', contigs = top_hits(full_zib_95_eb, alpha = 0.05)$Contig_name, show_points = T, plot_type = 'box')
Found 1 tophits for spikedTRUE at alpha = 0.05 using holm 
Warning: Removed 153 rows containing non-finite outside the scale range
(`stat_boxplot()`).

Still no FP as expected. However, eBayes priors were unnecessary for this case. Larger datasets tend to work okay with either weak or strong prior options. Faster to use and maybe more stable.

Small sample test

This is where ebp priors shine! Here we sample and take 16 spiked vs. 17 non spiked.

Subset data and prepare for simulations

Show code
sample_sz = 33

# hack to convert to function
n1 <- floor(sample_sz * 0.5) # Let's do a half split
n2 <- sample_sz - n1

set.seed(1988)
meta_subset <- bind_rows(
  meta_data %>% filter(spiked == TRUE) %>% slice_sample(n = n1),
  meta_data %>% filter(spiked == FALSE) %>% slice_sample(n = n2)
)

d_subset = d_p95[, meta_subset$run_accession]
d_subset <- filter_by_presence(d_subset, min_nonzero = ceiling(max(sample_sz*0.1, 2)))
Retained 550 rows after filtering
Show code
d_subset = modify_metadata(d_subset, meta_subset)
dim(d_subset)
[1] 550  33

Fit ZIB weak prior

Show code
save_path = "output_rds/prior_zib_small_weak.rds"

if(file.exists(save_path)){
  fit_zib_w <- readRDS(save_path)
} else {
  system.time({
    fit_zib_w <- glmZiBFit(d_subset,  design, nthreads=parallel::detectCores())
  })
  saveRDS(fit_zib_w, save_path)
}

head(top_hits(fit_zib_w))
Found 15 tophits for spikedTRUE at alpha = 0.05 using holm 
# A tibble: 6 × 10
  Contig_name Genome_file coefficient std_error  p_value p_adjust zi_coefficient
  <chr>       <chr>             <dbl>     <dbl>    <dbl>    <dbl>          <dbl>
1 UQDC010000… GCA_900539…     -0.213    0.0130  4.20e-60 2.31e-57         1.13  
2 HF995976.1… GCA_000432…     -0.0396   0.00343 8.07e-31 4.43e-28        -0.0240
3 NZ_JAHLPX0… GCF_018918…      0.421    0.0500  3.80e-17 2.08e-14        -0.366 
4 CAAFEQ0100… GCA_900761…     -0.505    0.0817  6.61e-10 3.61e- 7         1.13  
5 URBY010000… GCA_900546…     -0.0606   0.00987 8.16e-10 4.46e- 7        -1.14  
6 CAKSTI0100… GCA_934500…      0.118    0.0196  1.73e- 9 9.43e- 7        -0.0240
# ℹ 3 more variables: zi_std_error <dbl>, zi_p_value <dbl>, zi_p_adjust <dbl>
Show code
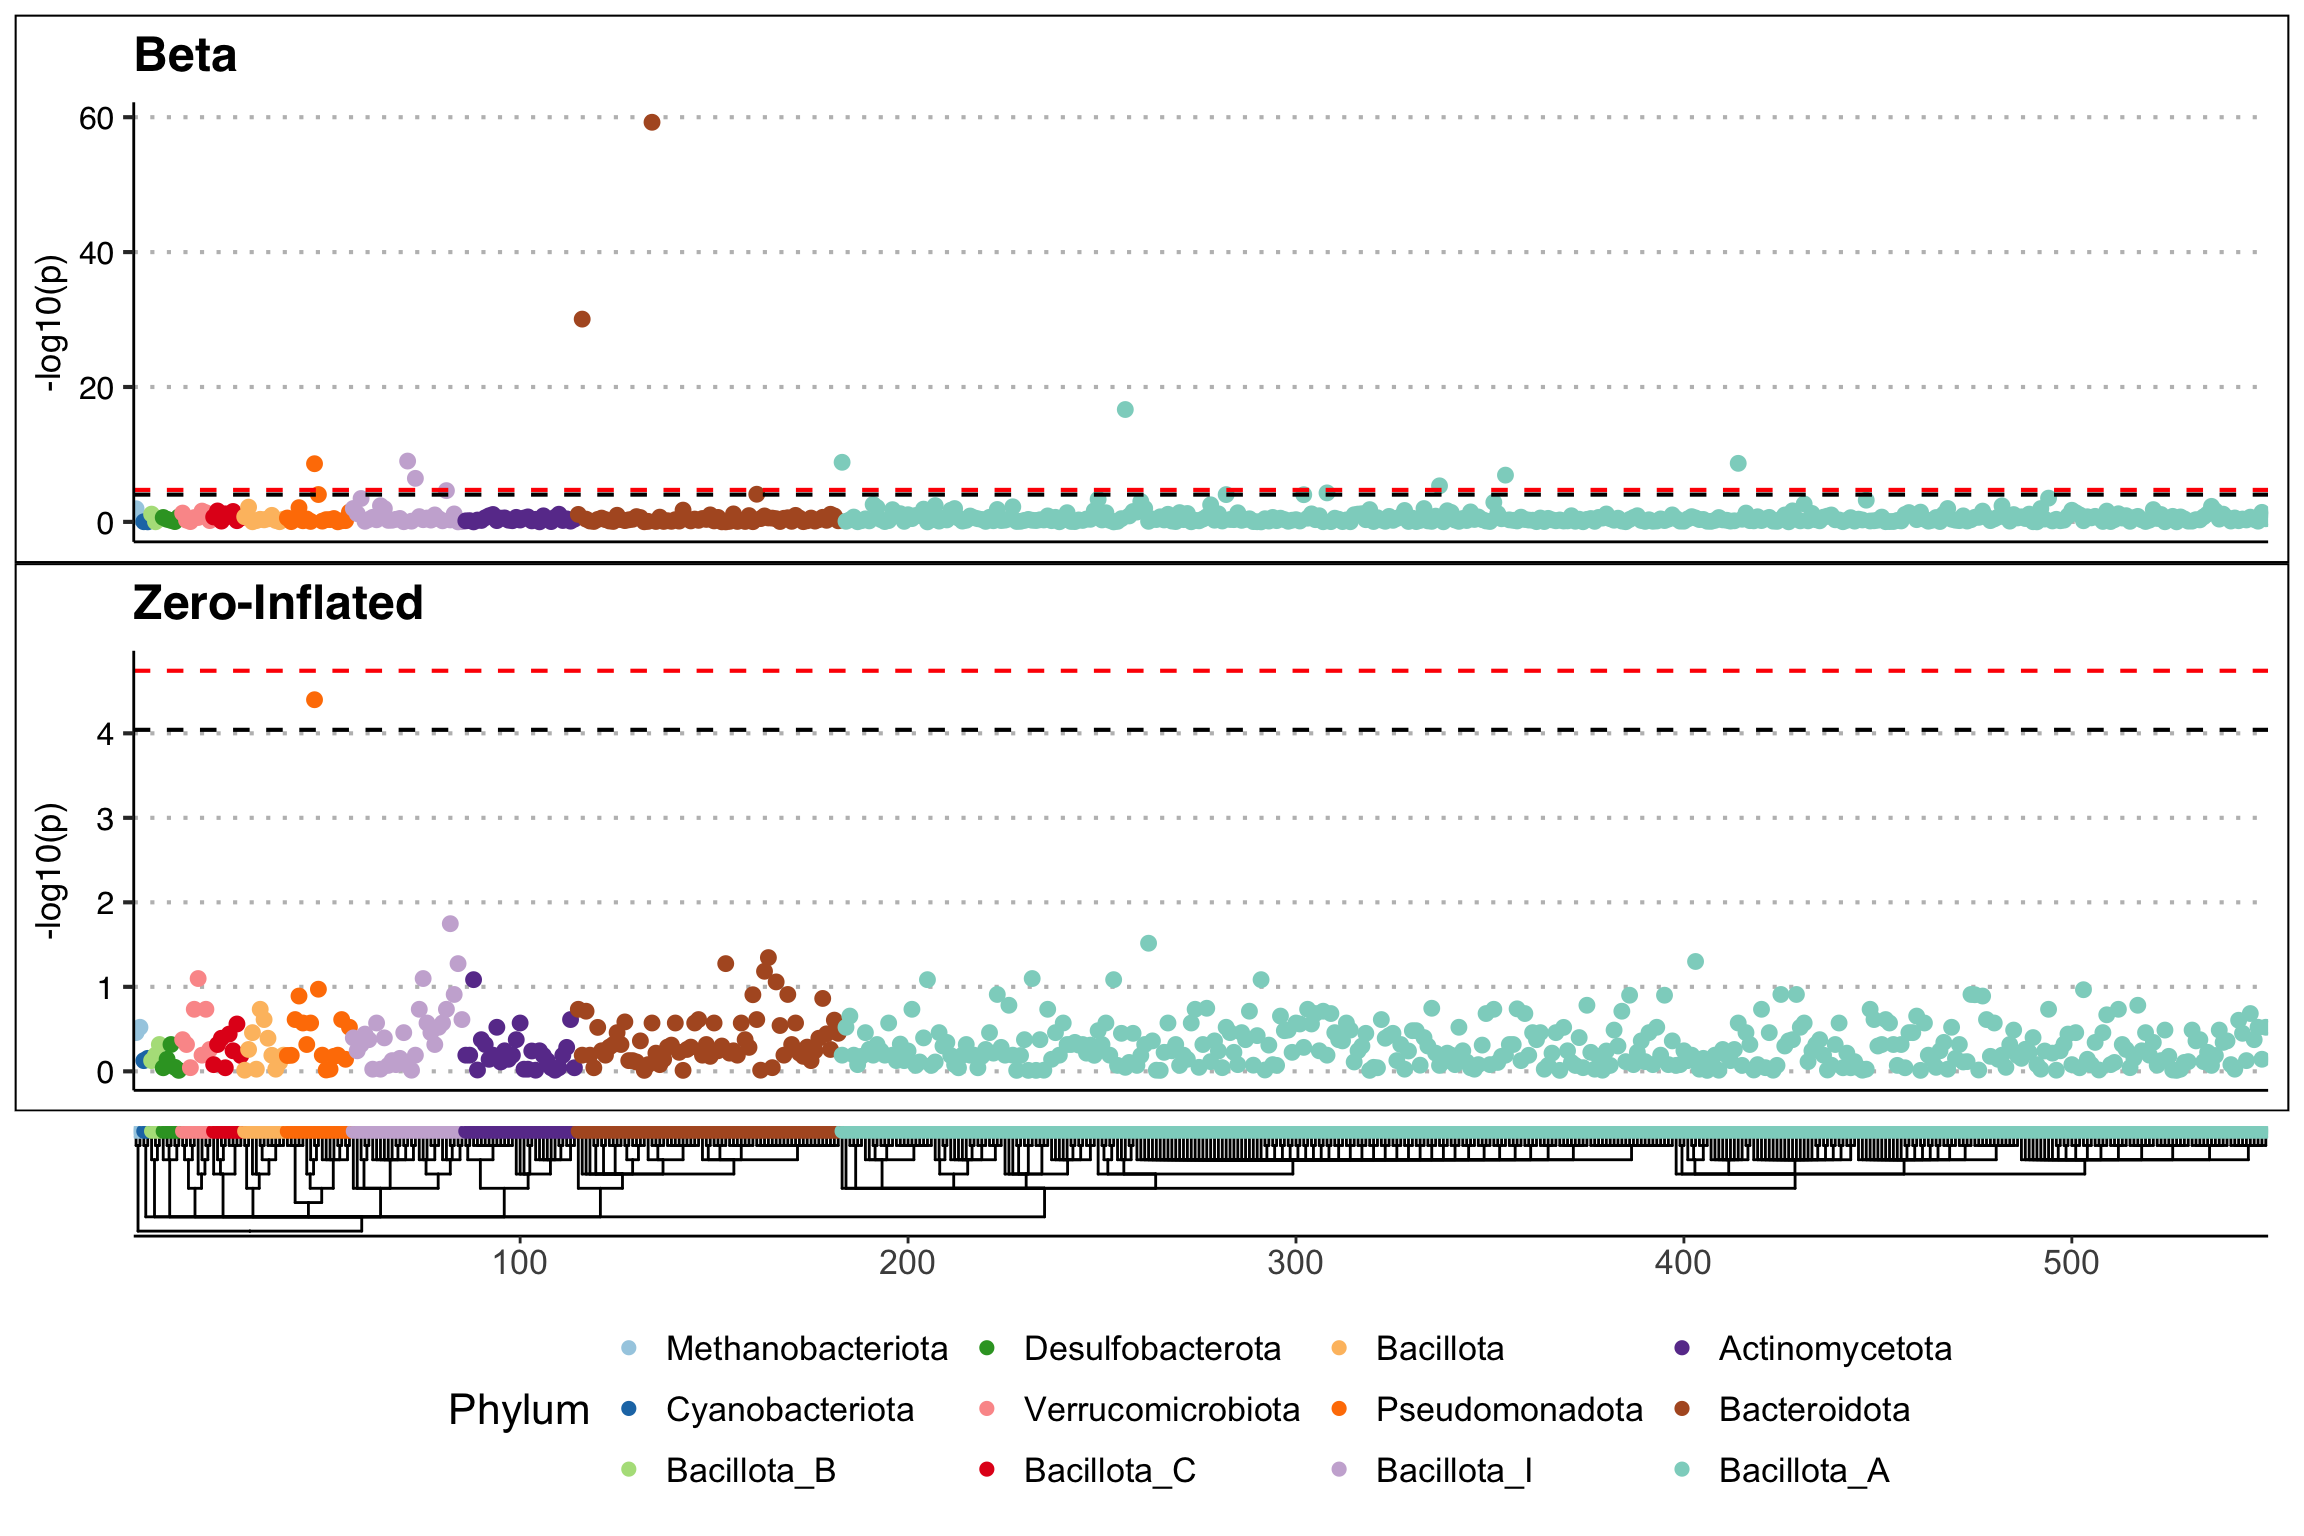
plot_manhattan(fit_zib_w, taxonomy = tax_95, aggregate_by_taxa = F)
! # Invaild edge matrix for <phylo>. A <tbl_df> is returned.

Show code
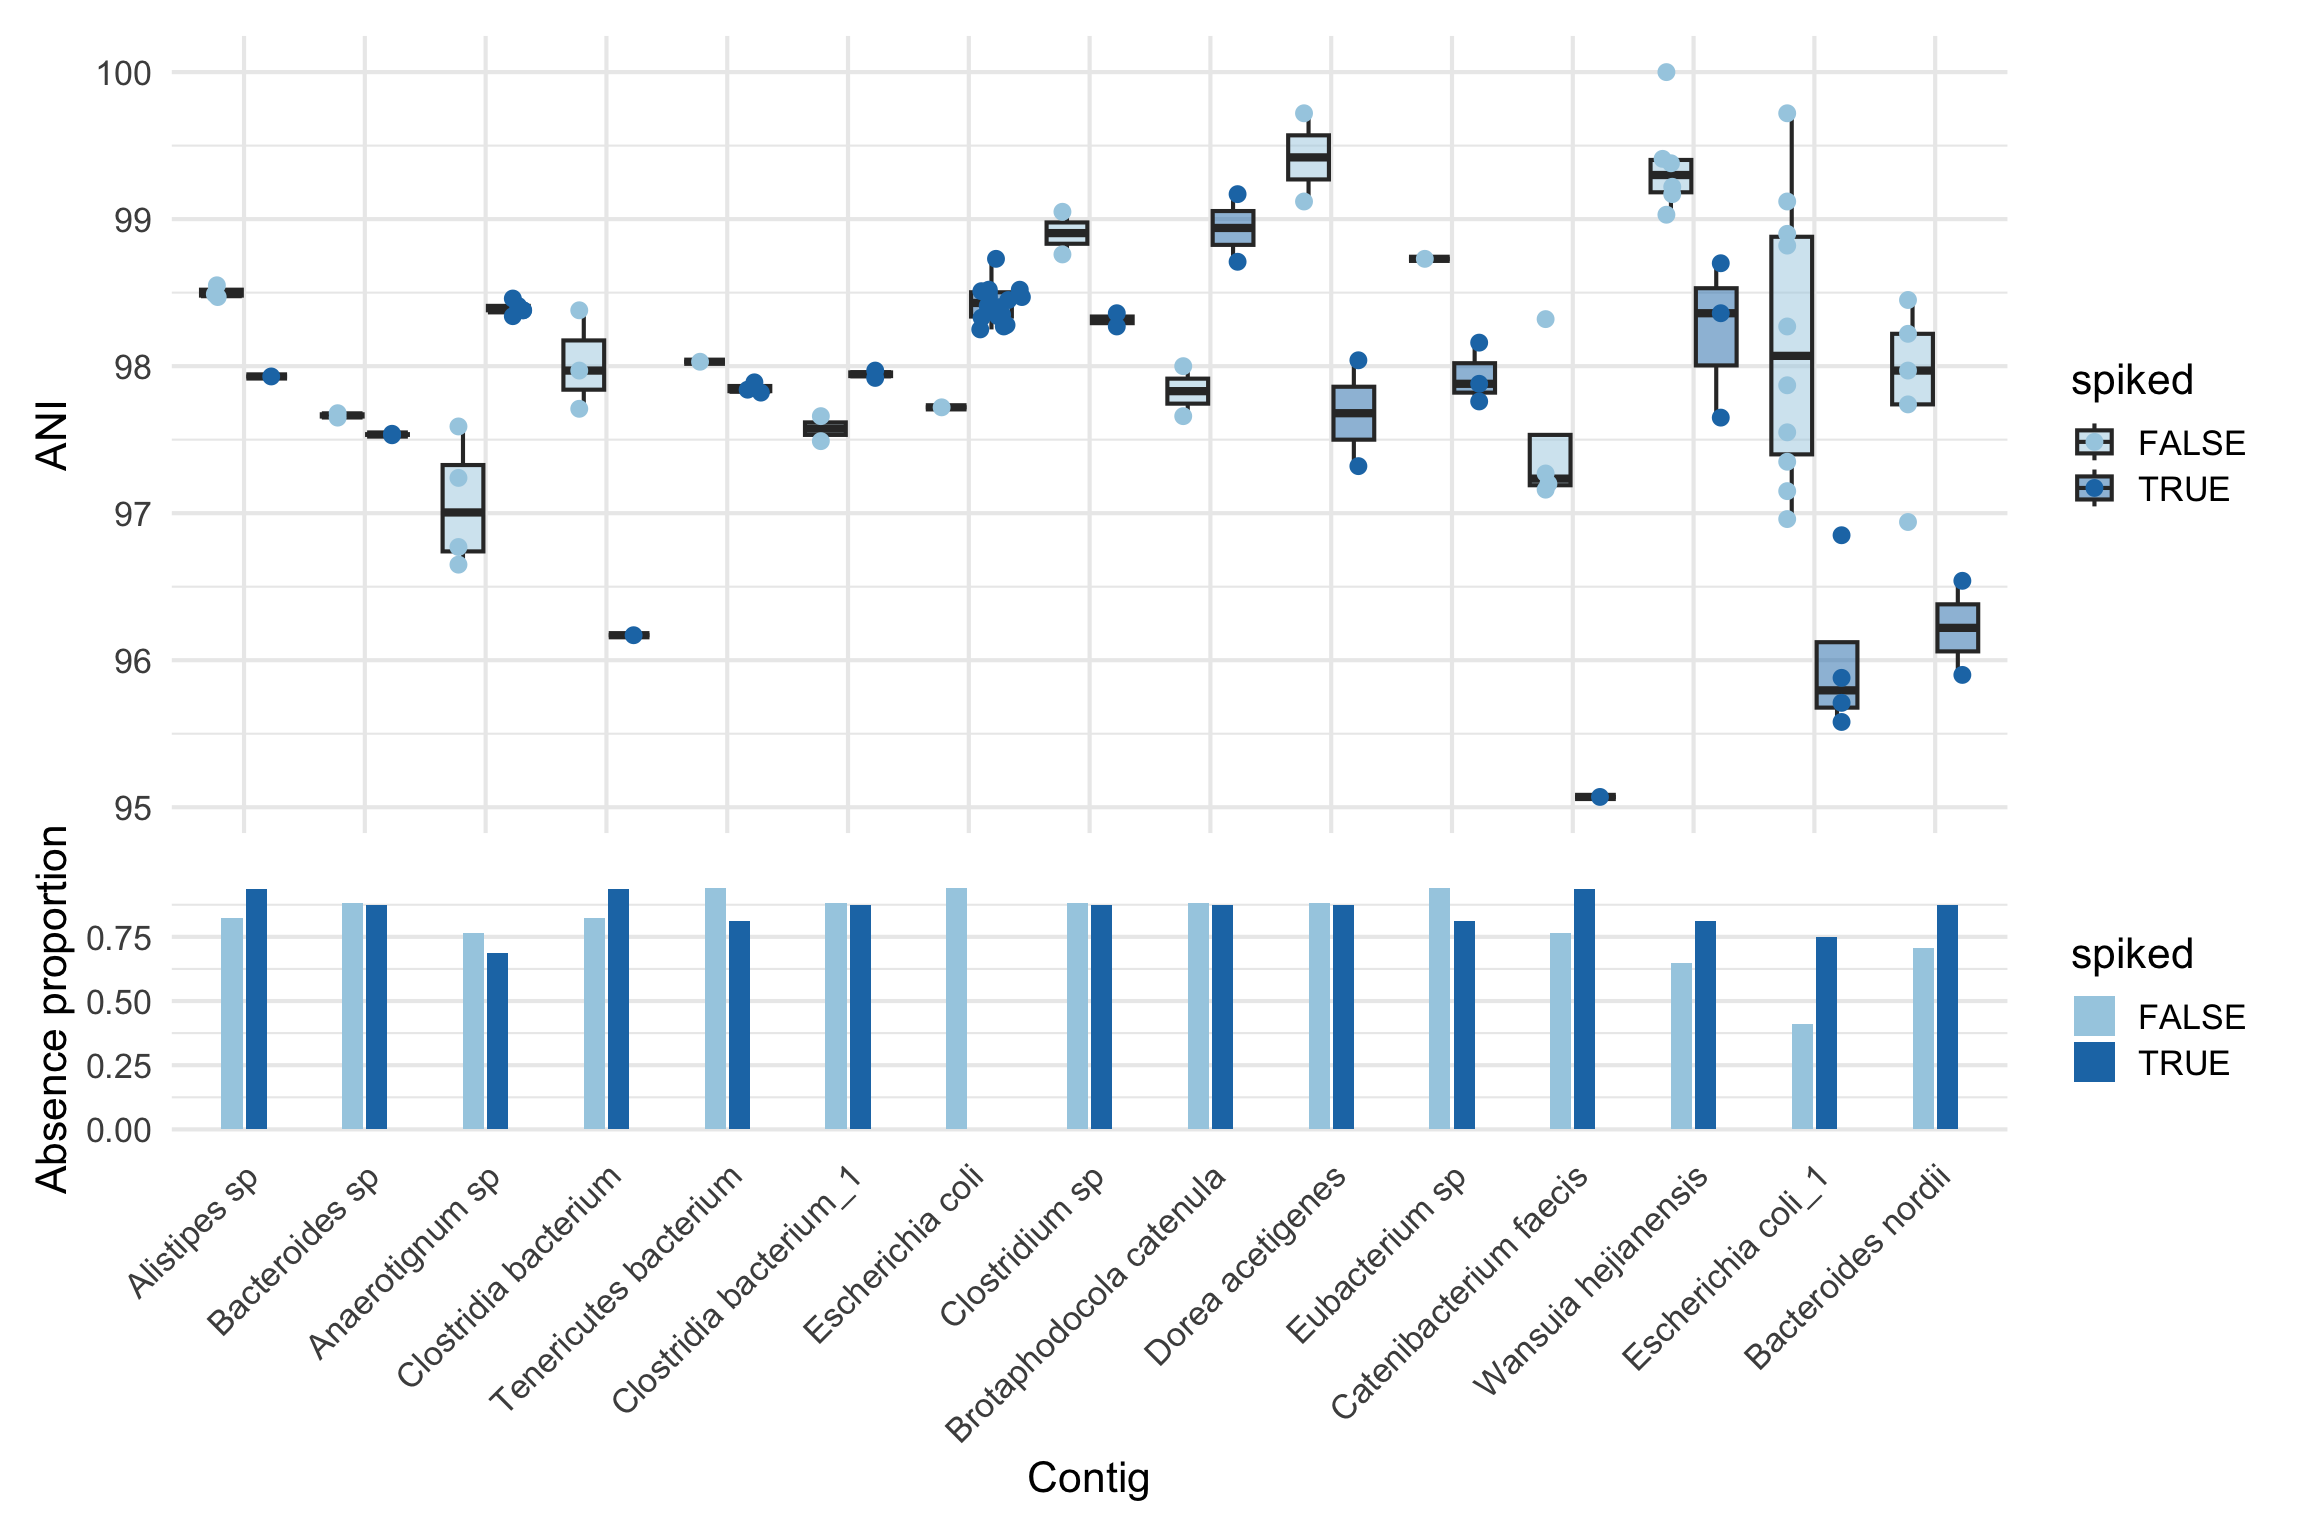
# Plenty of edge cases
plot_ani_dist(d_subset, phenotype = 'spiked', contigs = top_hits(fit_zib_w, alpha = 0.05)$Contig_name[1:15], show_points = T, plot_type = 'box')
Found 15 tophits for spikedTRUE at alpha = 0.05 using holm 
Warning: Removed 398 rows containing non-finite outside the scale range
(`stat_boxplot()`).

Fit ZIB strong prior

Show code
save_path = "output_rds/prior_zib_small_strong.rds"

if(file.exists(save_path)){
  fit_zib_s <- readRDS(save_path)
} else {
  system.time({
    fit_zib_s <- glmZiBFit(d_subset,  design, nthreads=parallel::detectCores(), MAP_prior = "preset_strong")
  })
  saveRDS(fit_zib_s, save_path)
}

head(top_hits(fit_zib_s))
Found 15 tophits for spikedTRUE at alpha = 0.05 using holm 
# A tibble: 6 × 10
  Contig_name Genome_file coefficient std_error  p_value p_adjust zi_coefficient
  <chr>       <chr>             <dbl>     <dbl>    <dbl>    <dbl>          <dbl>
1 UQDC010000… GCA_900539…     -0.212    0.0130  5.61e-60 3.08e-57         0.762 
2 HF995976.1… GCA_000432…     -0.0396   0.00343 8.64e-31 4.74e-28         0.307 
3 NZ_JAHLPX0… GCF_018918…      0.423    0.0499  2.19e-17 1.20e-14        -0.0757
4 URBY010000… GCA_900546…     -0.0604   0.00987 9.62e-10 5.26e- 7        -0.122 
5 CAKSTI0100… GCA_934500…      0.118    0.0196  1.46e- 9 7.99e- 7         0.307 
6 CAAFEQ0100… GCA_900761…     -0.494    0.0824  2.09e- 9 1.14e- 6         0.762 
# ℹ 3 more variables: zi_std_error <dbl>, zi_p_value <dbl>, zi_p_adjust <dbl>
Show code
plot_manhattan(fit_zib_s, taxonomy = tax_95, aggregate_by_taxa = F)
! # Invaild edge matrix for <phylo>. A <tbl_df> is returned.

Show code
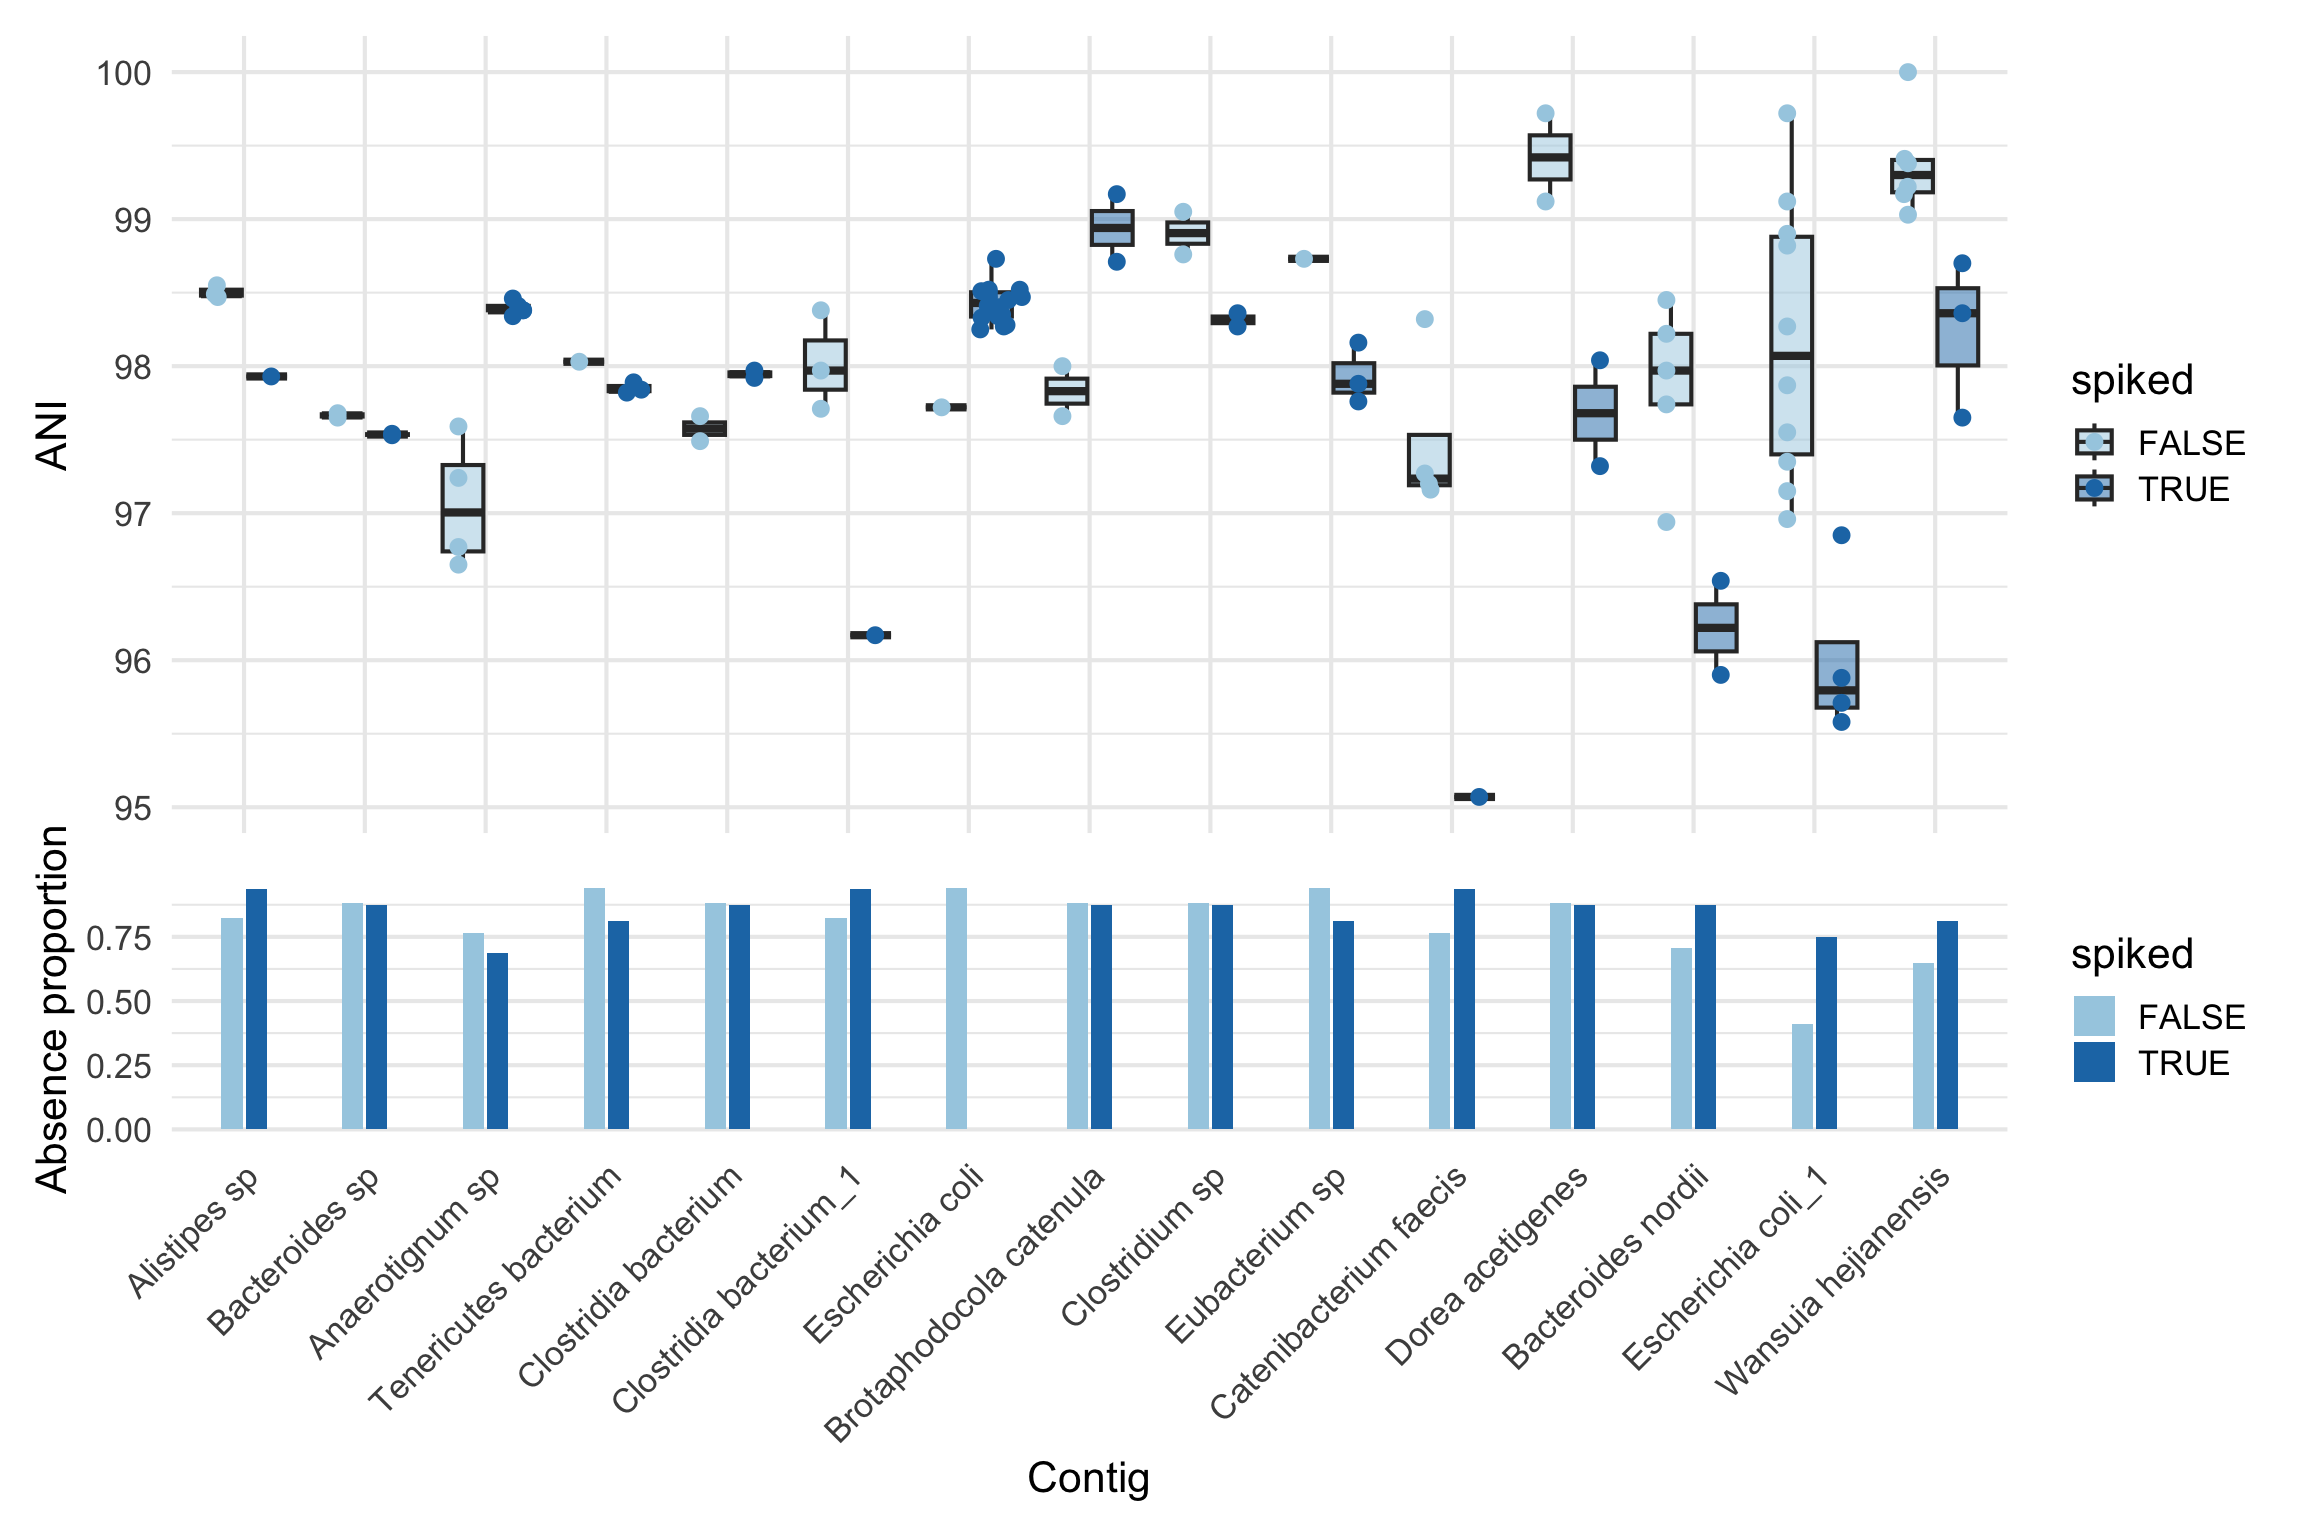
# Still edge cases
plot_ani_dist(d_subset, phenotype = 'spiked', contigs = top_hits(fit_zib_s, alpha = 0.05)$Contig_name,show_points = T, plot_type = 'box')
Found 15 tophits for spikedTRUE at alpha = 0.05 using holm 
Warning: Removed 398 rows containing non-finite outside the scale range
(`stat_boxplot()`).

Compare the two priors

Show code
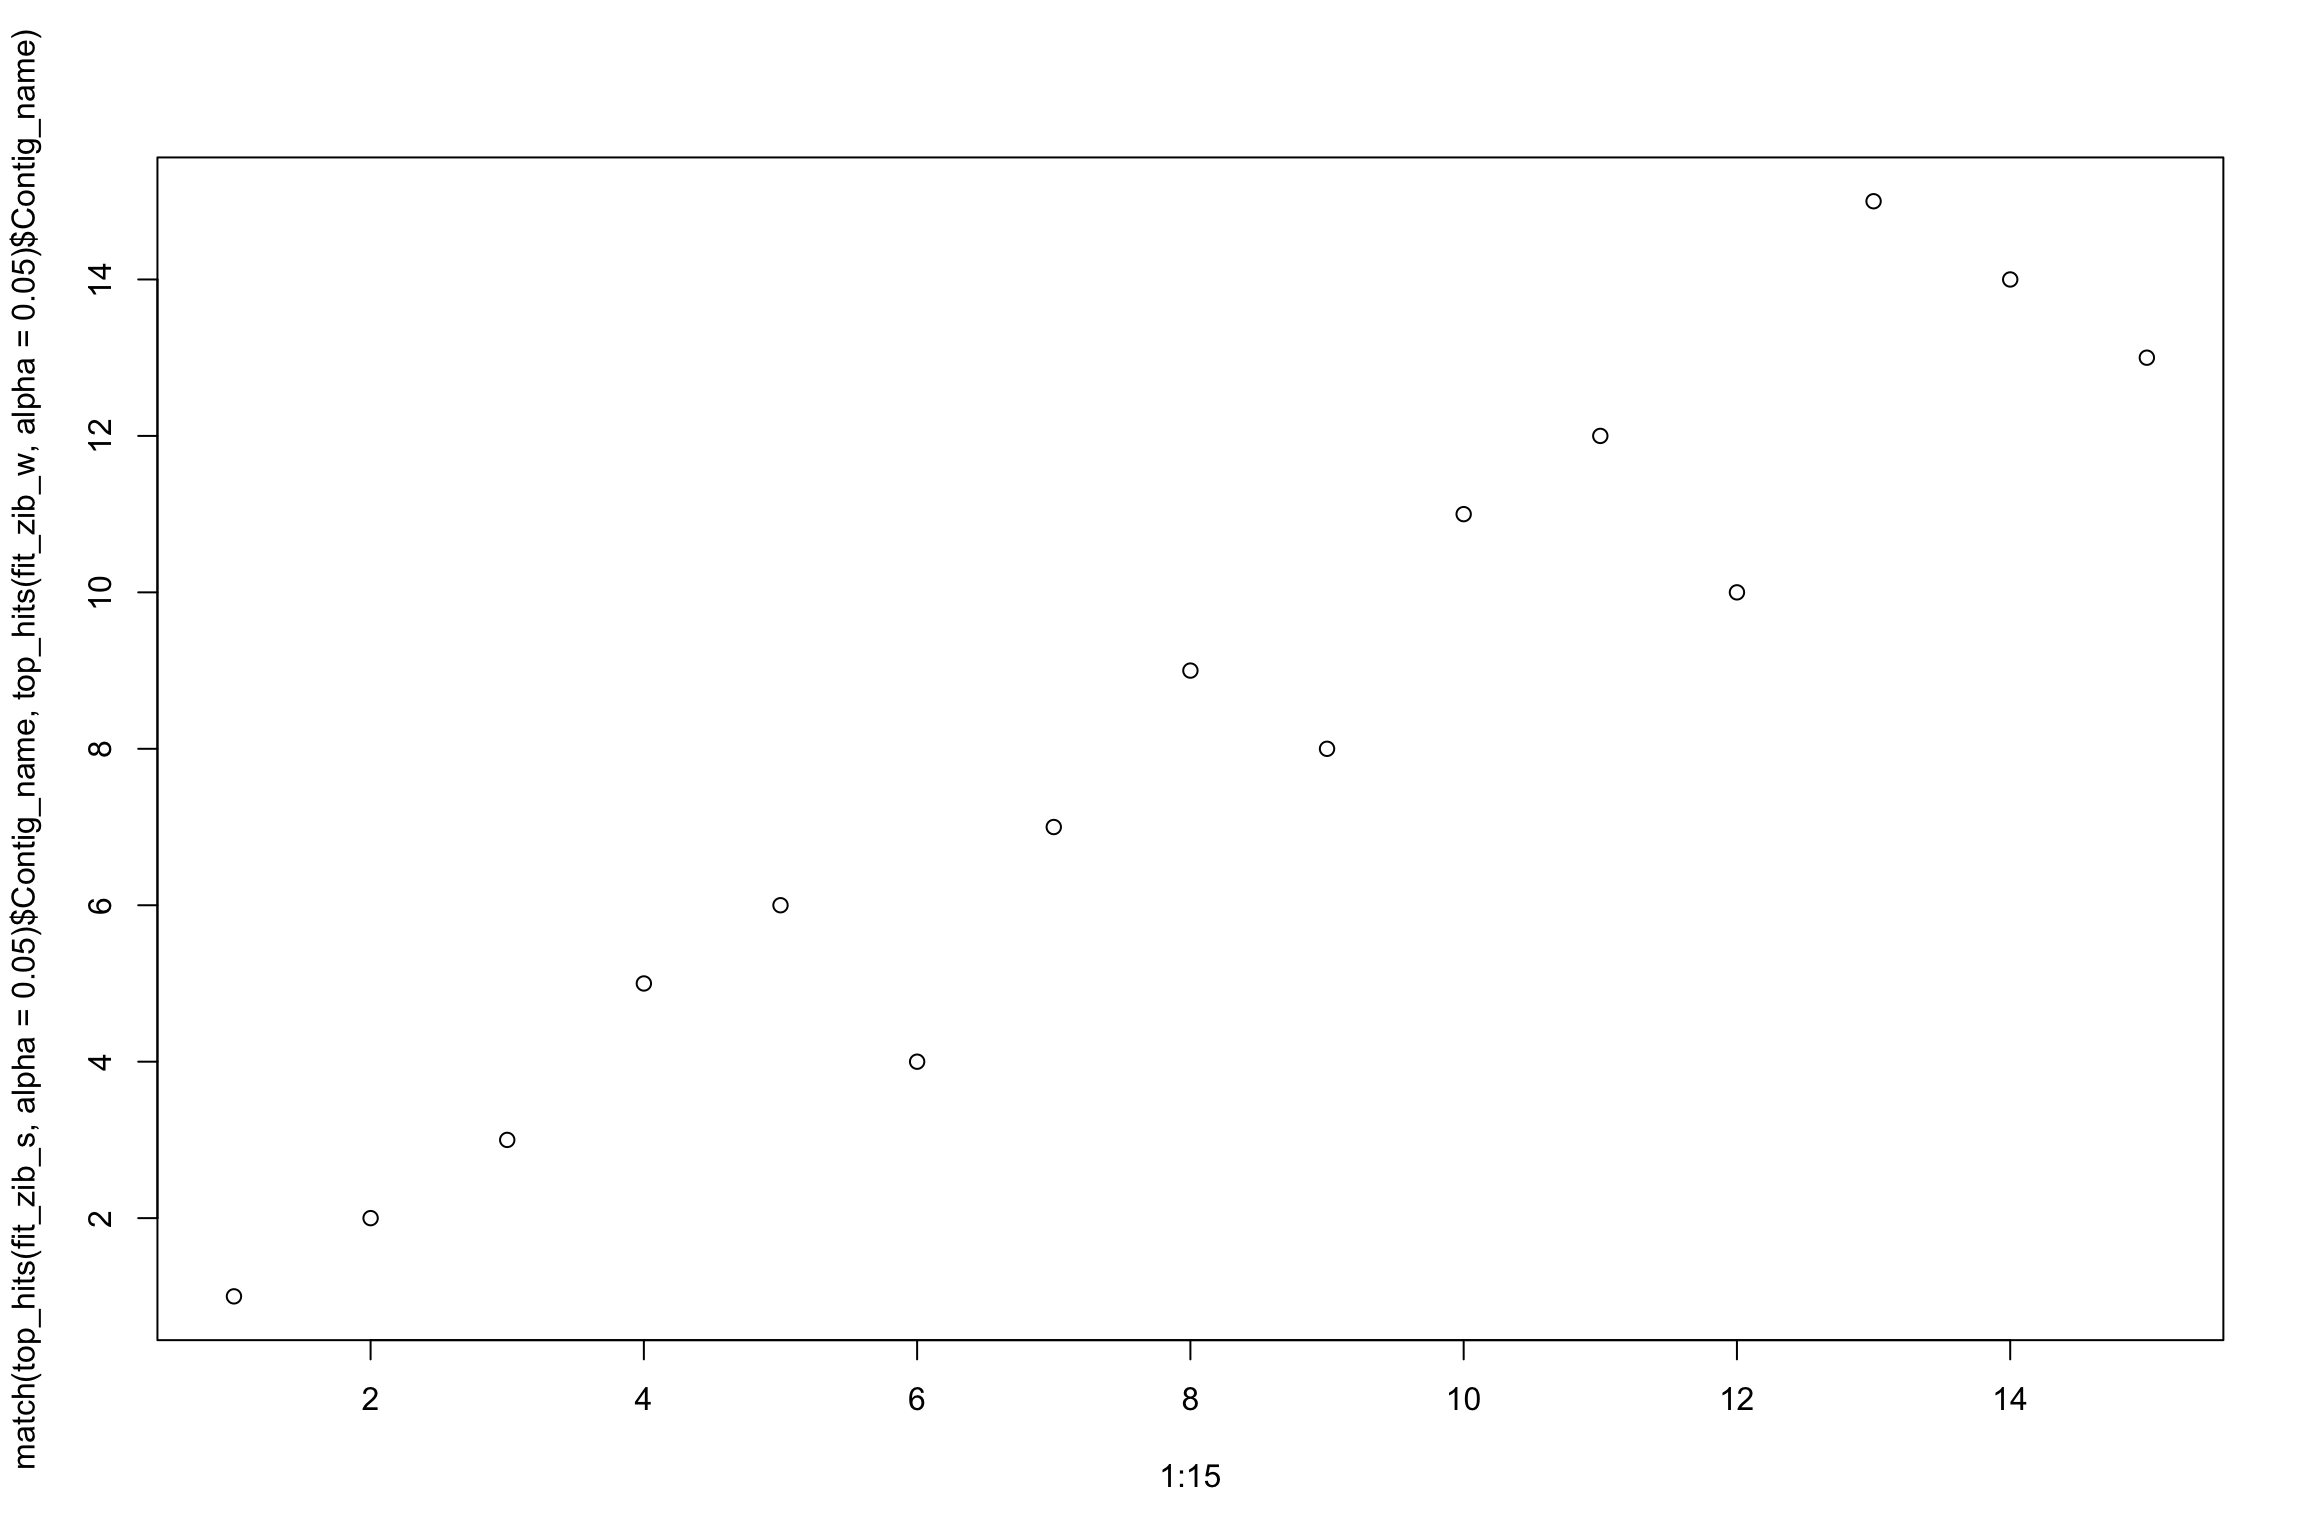
# At a strong multiple testing correction, both priors pick the same edge cases, but in a different order
all(top_hits(fit_zib_s, alpha = 0.05)$Contig_name %in% top_hits(fit_zib_w, alpha = 0.05)$Contig_name)
Found 15 tophits for spikedTRUE at alpha = 0.05 using holm 
[1] TRUE
Show code
plot(1:15, match(top_hits(fit_zib_s, alpha = 0.05)$Contig_name, top_hits(fit_zib_w, alpha = 0.05)$Contig_name))
Found 15 tophits for spikedTRUE at alpha = 0.05 using holm 

Show code
# But if we relax the multple correction method to BH, the weak prior has more false positives than the strong prior
length(top_hits(fit_zib_w, alpha = 0.05, method = 'BH')$Contig_name)
Found 28 tophits for spikedTRUE at alpha = 0.05 using BH 
[1] 28
Show code
length(top_hits(fit_zib_s, alpha = 0.05, method = 'BH')$Contig_name) 
Found 22 tophits for spikedTRUE at alpha = 0.05 using BH 
[1] 22
Show code
# this is as expected

Both methods are not great for the small sample size.

Fit empirical bayes prior

Show code
ebp = compute_eb_priors(d_subset, design = design, nthreads = parallel::detectCores(), low_cutoff = 0, high_cutoff = Inf, progress = FALSE)
ebp
strainspy_priors object
Method:  empirical 

Priors:
 - fixef : 1 coefficients
 - fixef_zi : 1 coefficients

Example priors:
           prior    class       coef
1 normal(0,0.23)    fixef spikedTRUE
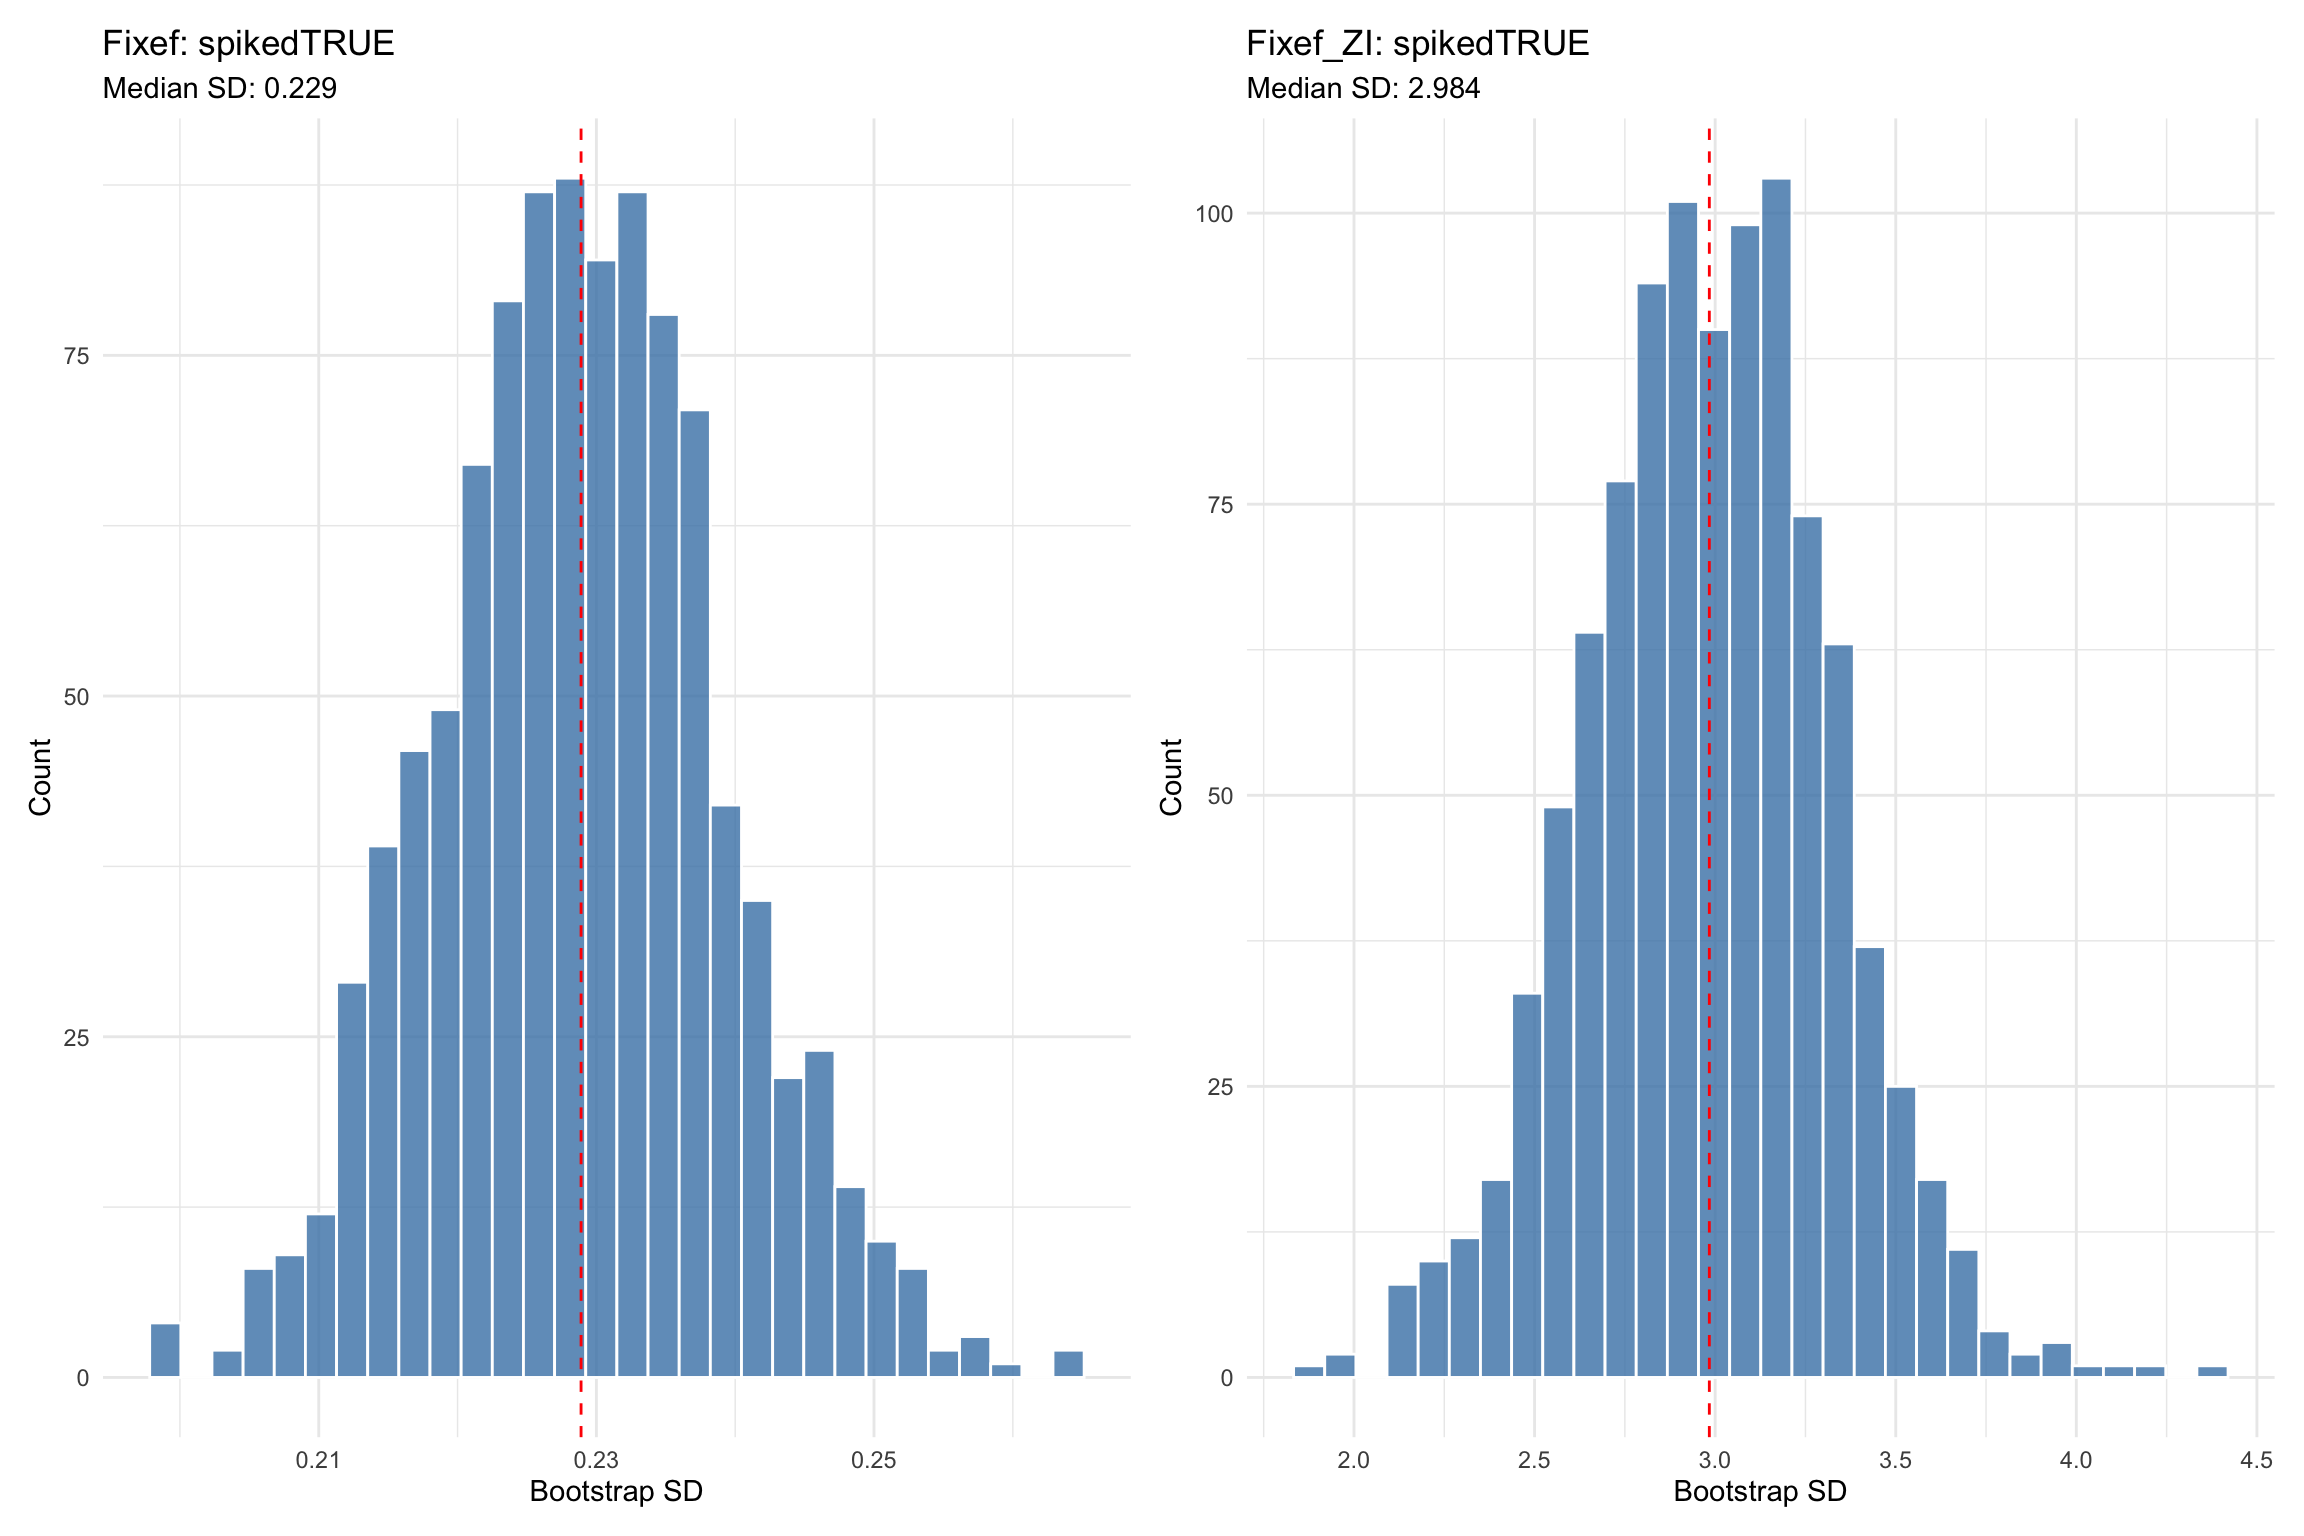
2 normal(0,2.98) fixef_zi spikedTRUE

Bootstrap summary for fixef priors:
   Min. 1st Qu.  Median    Mean 3rd Qu.    Max. 
 0.1989  0.2289  0.2575  0.2535  0.2782  0.3023 

Bootstrap summary for fixef_zi priors:
   Min. 1st Qu.  Median    Mean 3rd Qu.    Max. 
  1.118   2.193   2.588   2.580   3.000   4.377 

Warning: 2 fixef prior(s) may be too strong (SD < 1), and 0 may be too weak (SD > 5).
Show code
plot_prior_bootstrap(ebp, "spikedTRUE")

Show code
save_path = "output_rds/prior_zib_small_ebp.rds"

if(file.exists(save_path)){
  fit_zib_eb <- readRDS(save_path)
} else {
  system.time({
    fit_zib_eb <- glmZiBFit(d_subset,  design, nthreads=parallel::detectCores(), MAP_prior = ebp)
  })
  saveRDS(fit_zib_eb, save_path)
}


head(top_hits(fit_zib_eb))
Found 1 tophits for spikedTRUE at alpha = 0.05 using holm 
# A tibble: 1 × 10
  Contig_name  Genome_file coefficient std_error p_value p_adjust zi_coefficient
  <chr>        <chr>             <dbl>     <dbl>   <dbl>    <dbl>          <dbl>
1 NZ_AEZI0200… GCF_000194…       0.406     0.223  0.0691        1          -5.25
# ℹ 3 more variables: zi_std_error <dbl>, zi_p_value <dbl>, zi_p_adjust <dbl>
Show code
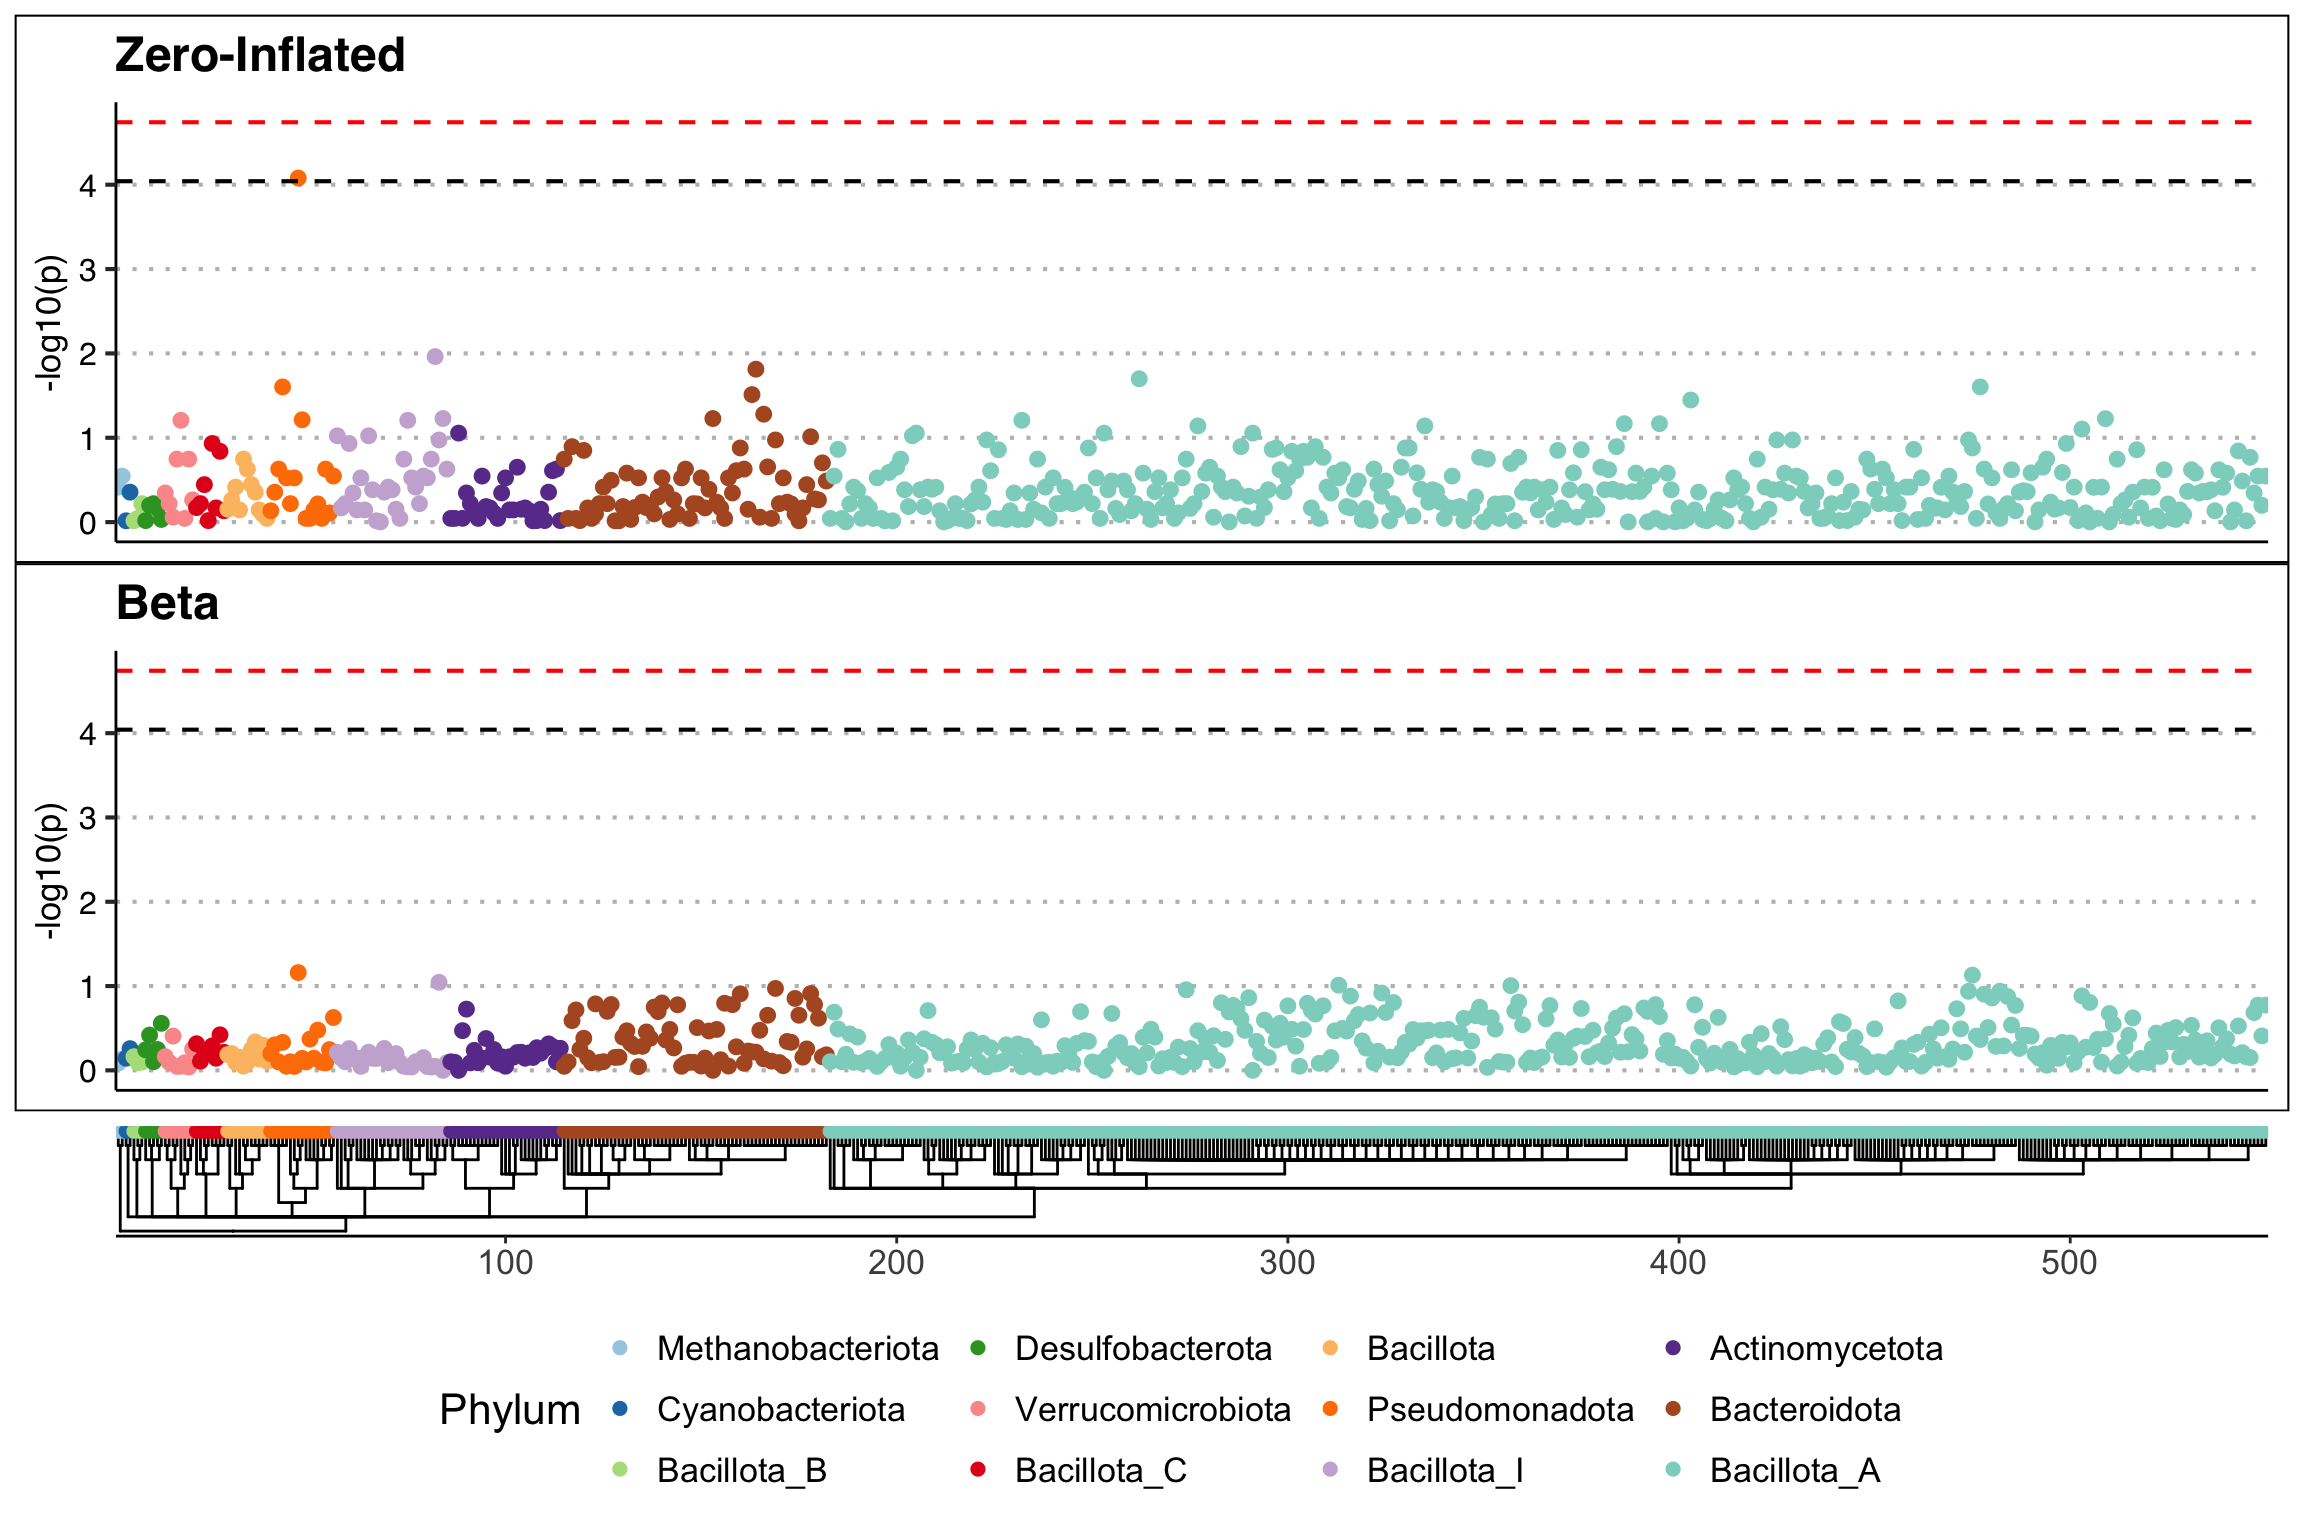
plot_manhattan(fit_zib_eb, taxonomy = tax_95, aggregate_by_taxa = F)
! # Invaild edge matrix for <phylo>. A <tbl_df> is returned.

Show code
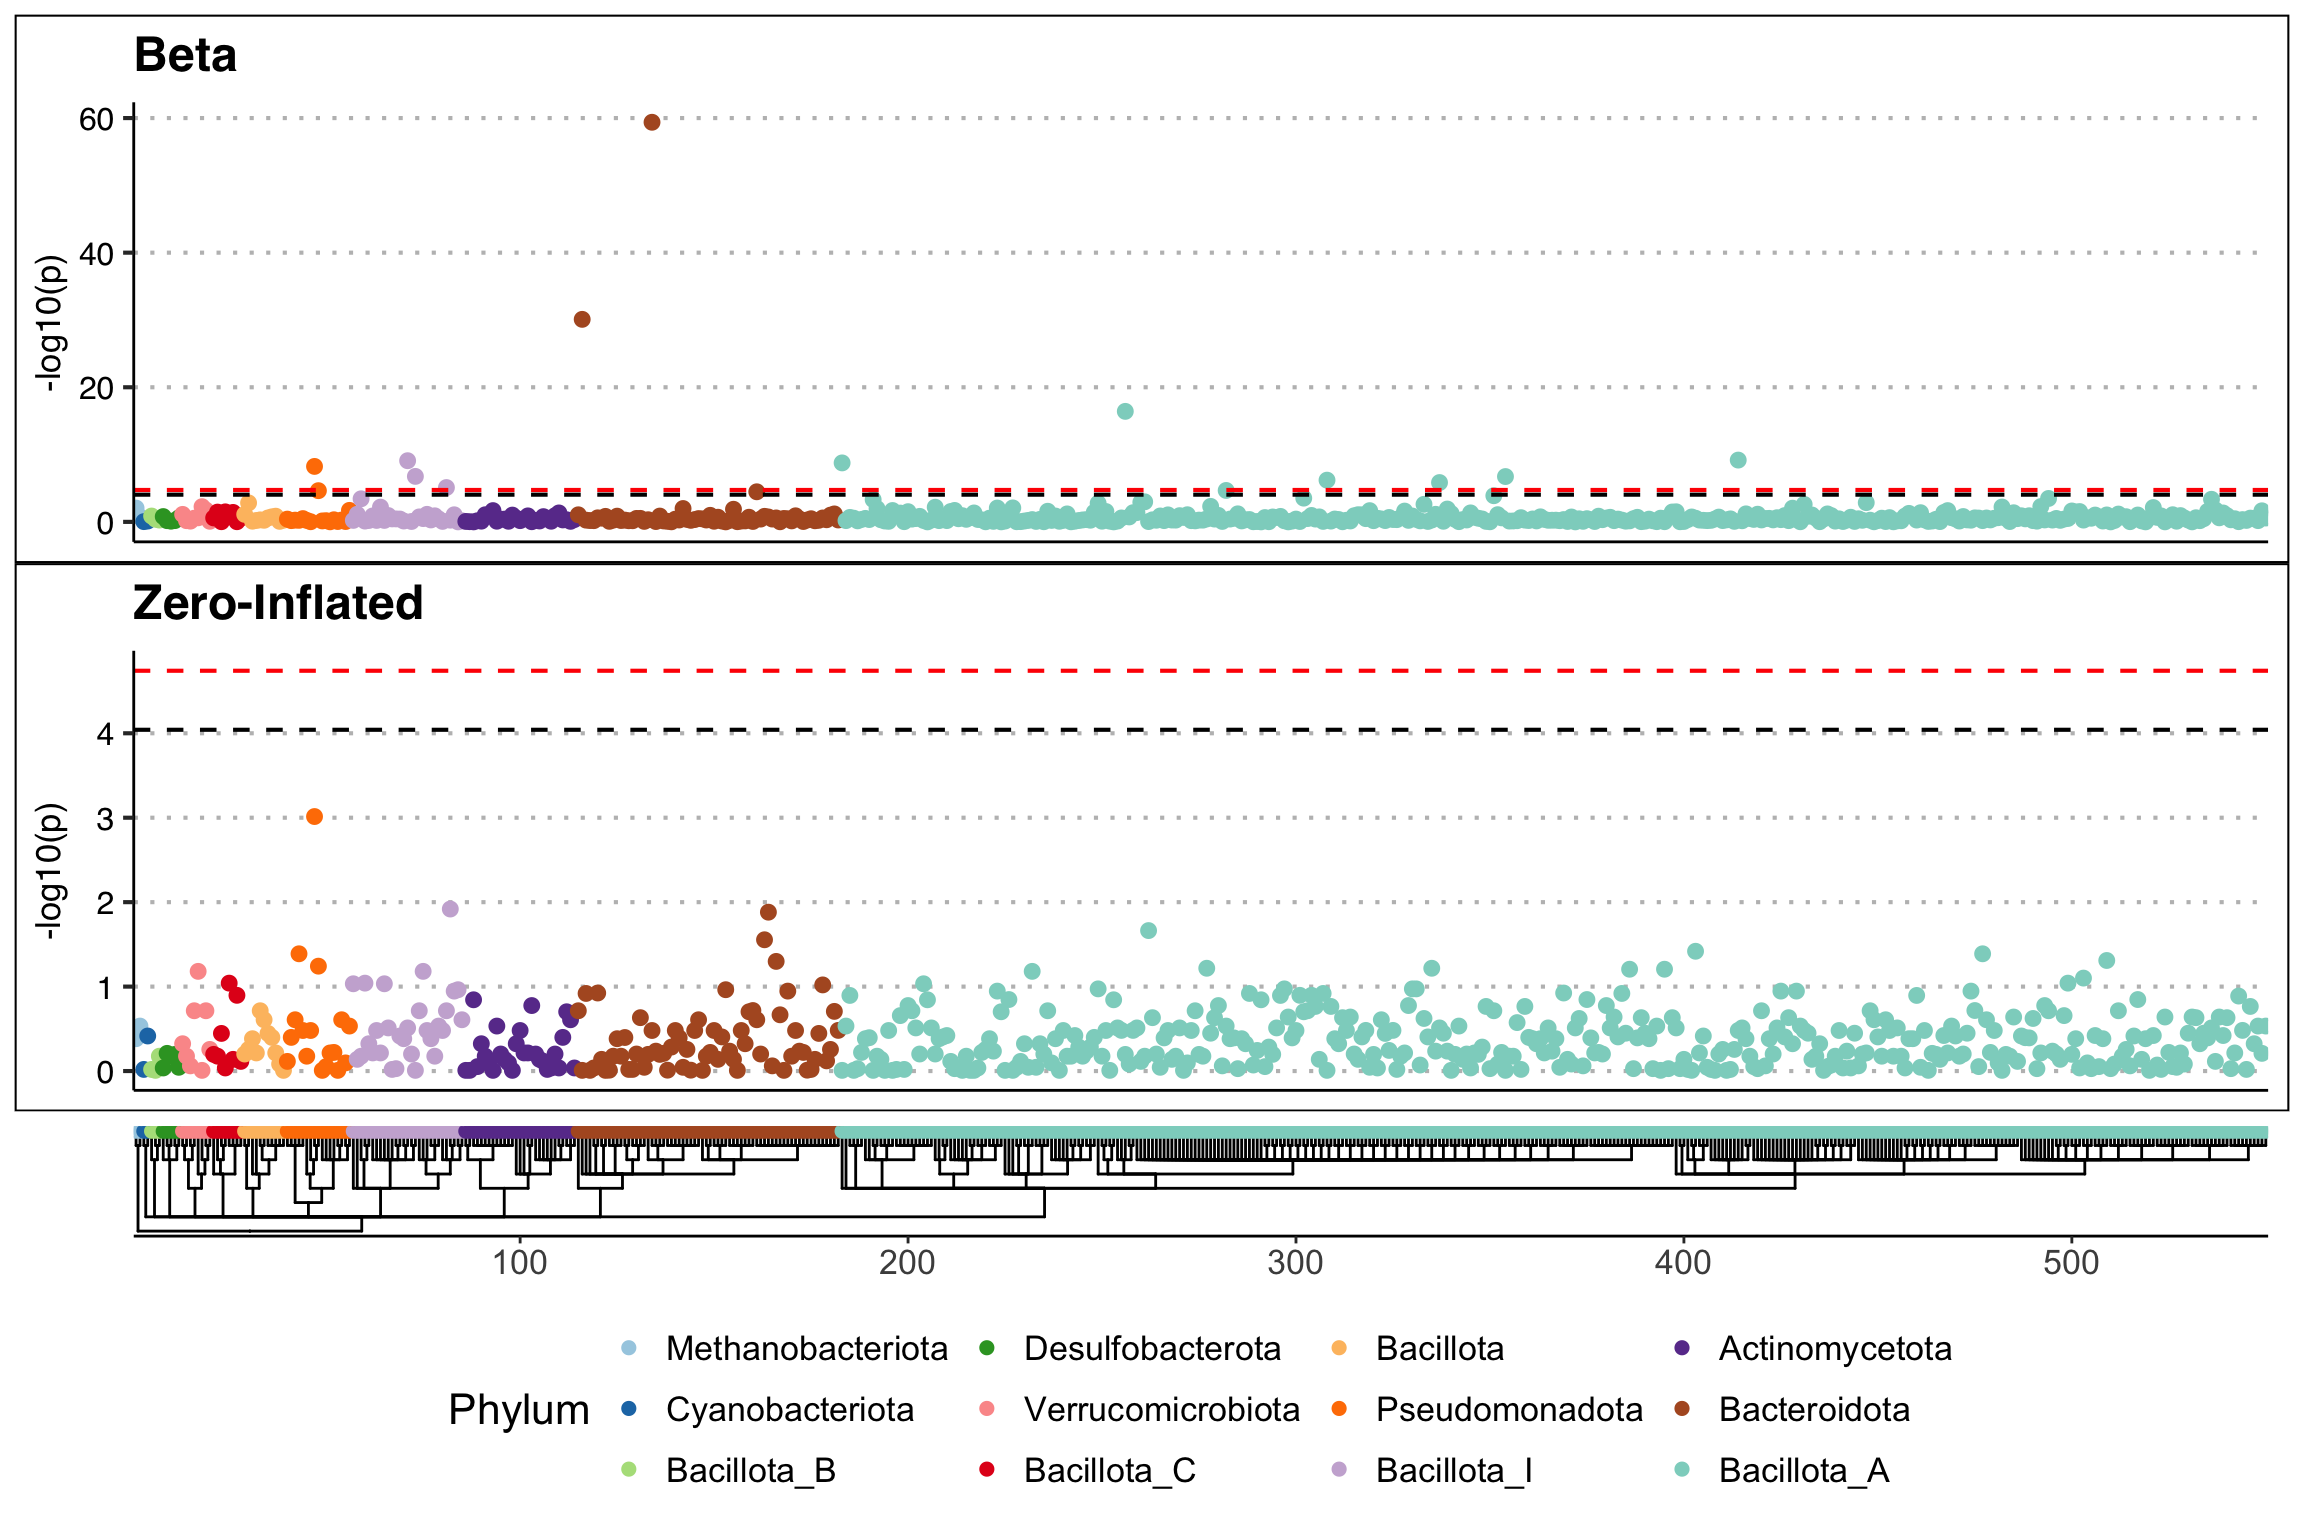
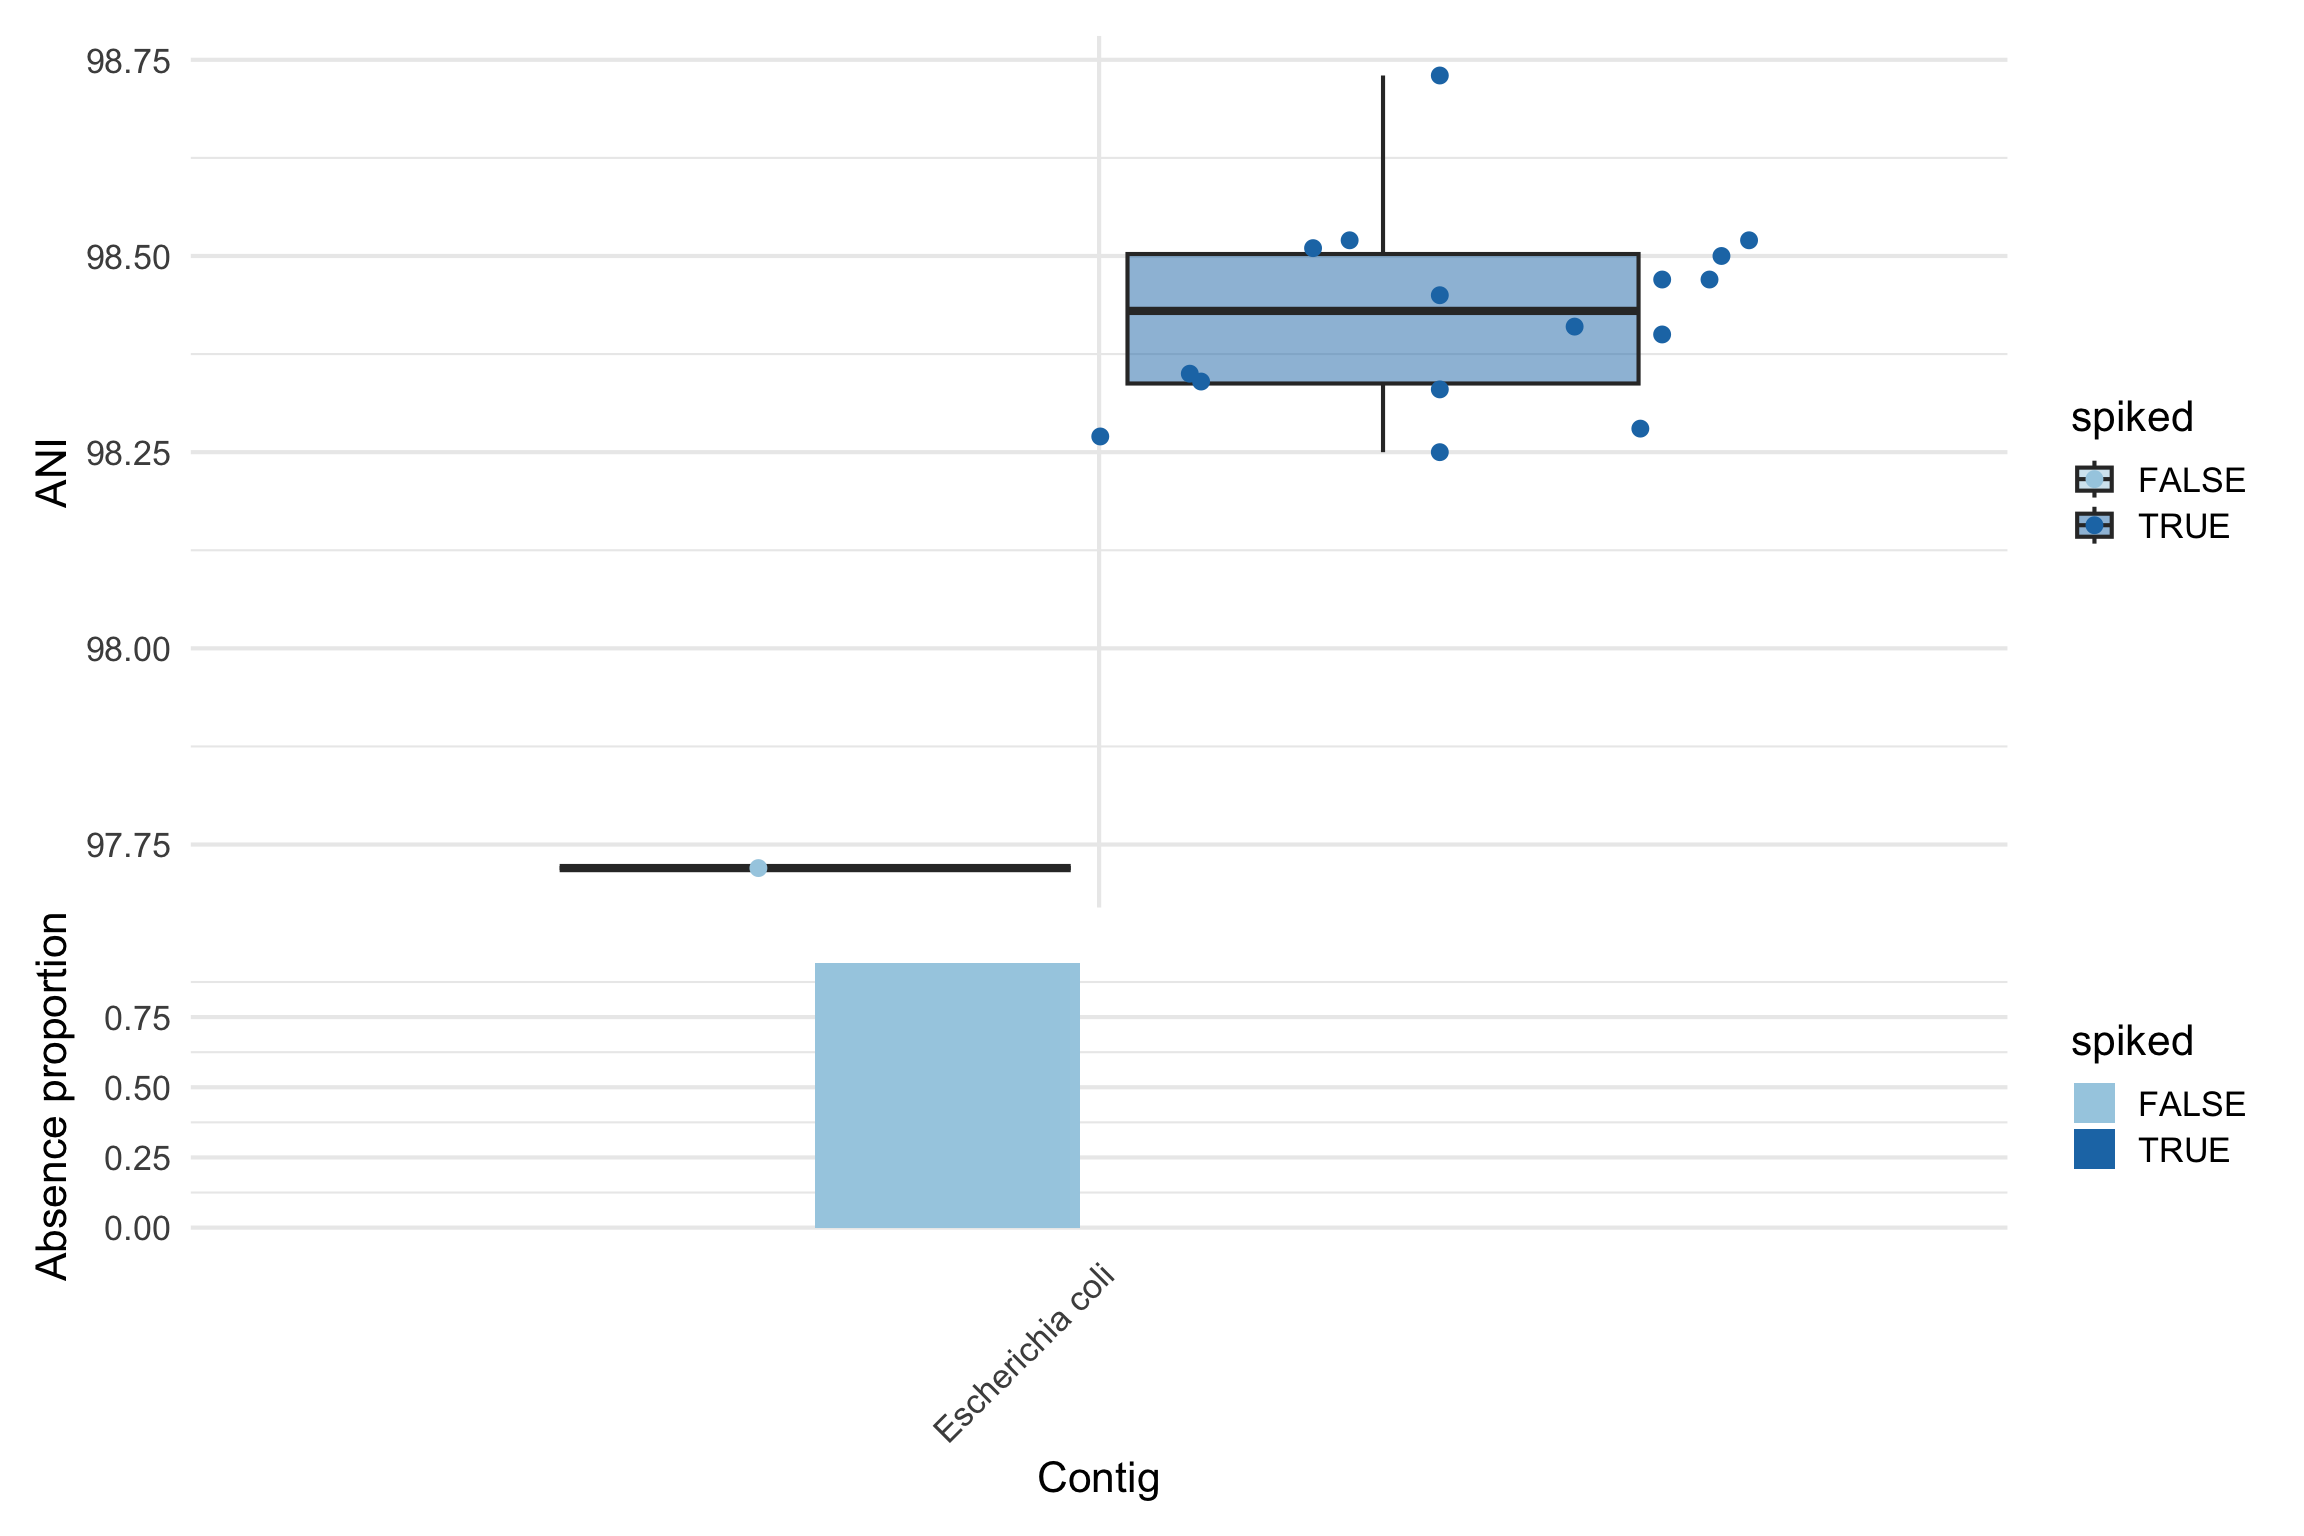
# We only have the ZI signal for E. coli now! 

# valid case
plot_ani_dist(d_subset, phenotype = 'spiked', contigs = top_hits(fit_zib_eb, alpha = 0.05)$Contig_name, show_points = T, plot_type = 'box')
Found 1 tophits for spikedTRUE at alpha = 0.05 using holm 
Warning: Removed 16 rows containing non-finite outside the scale range
(`stat_boxplot()`).
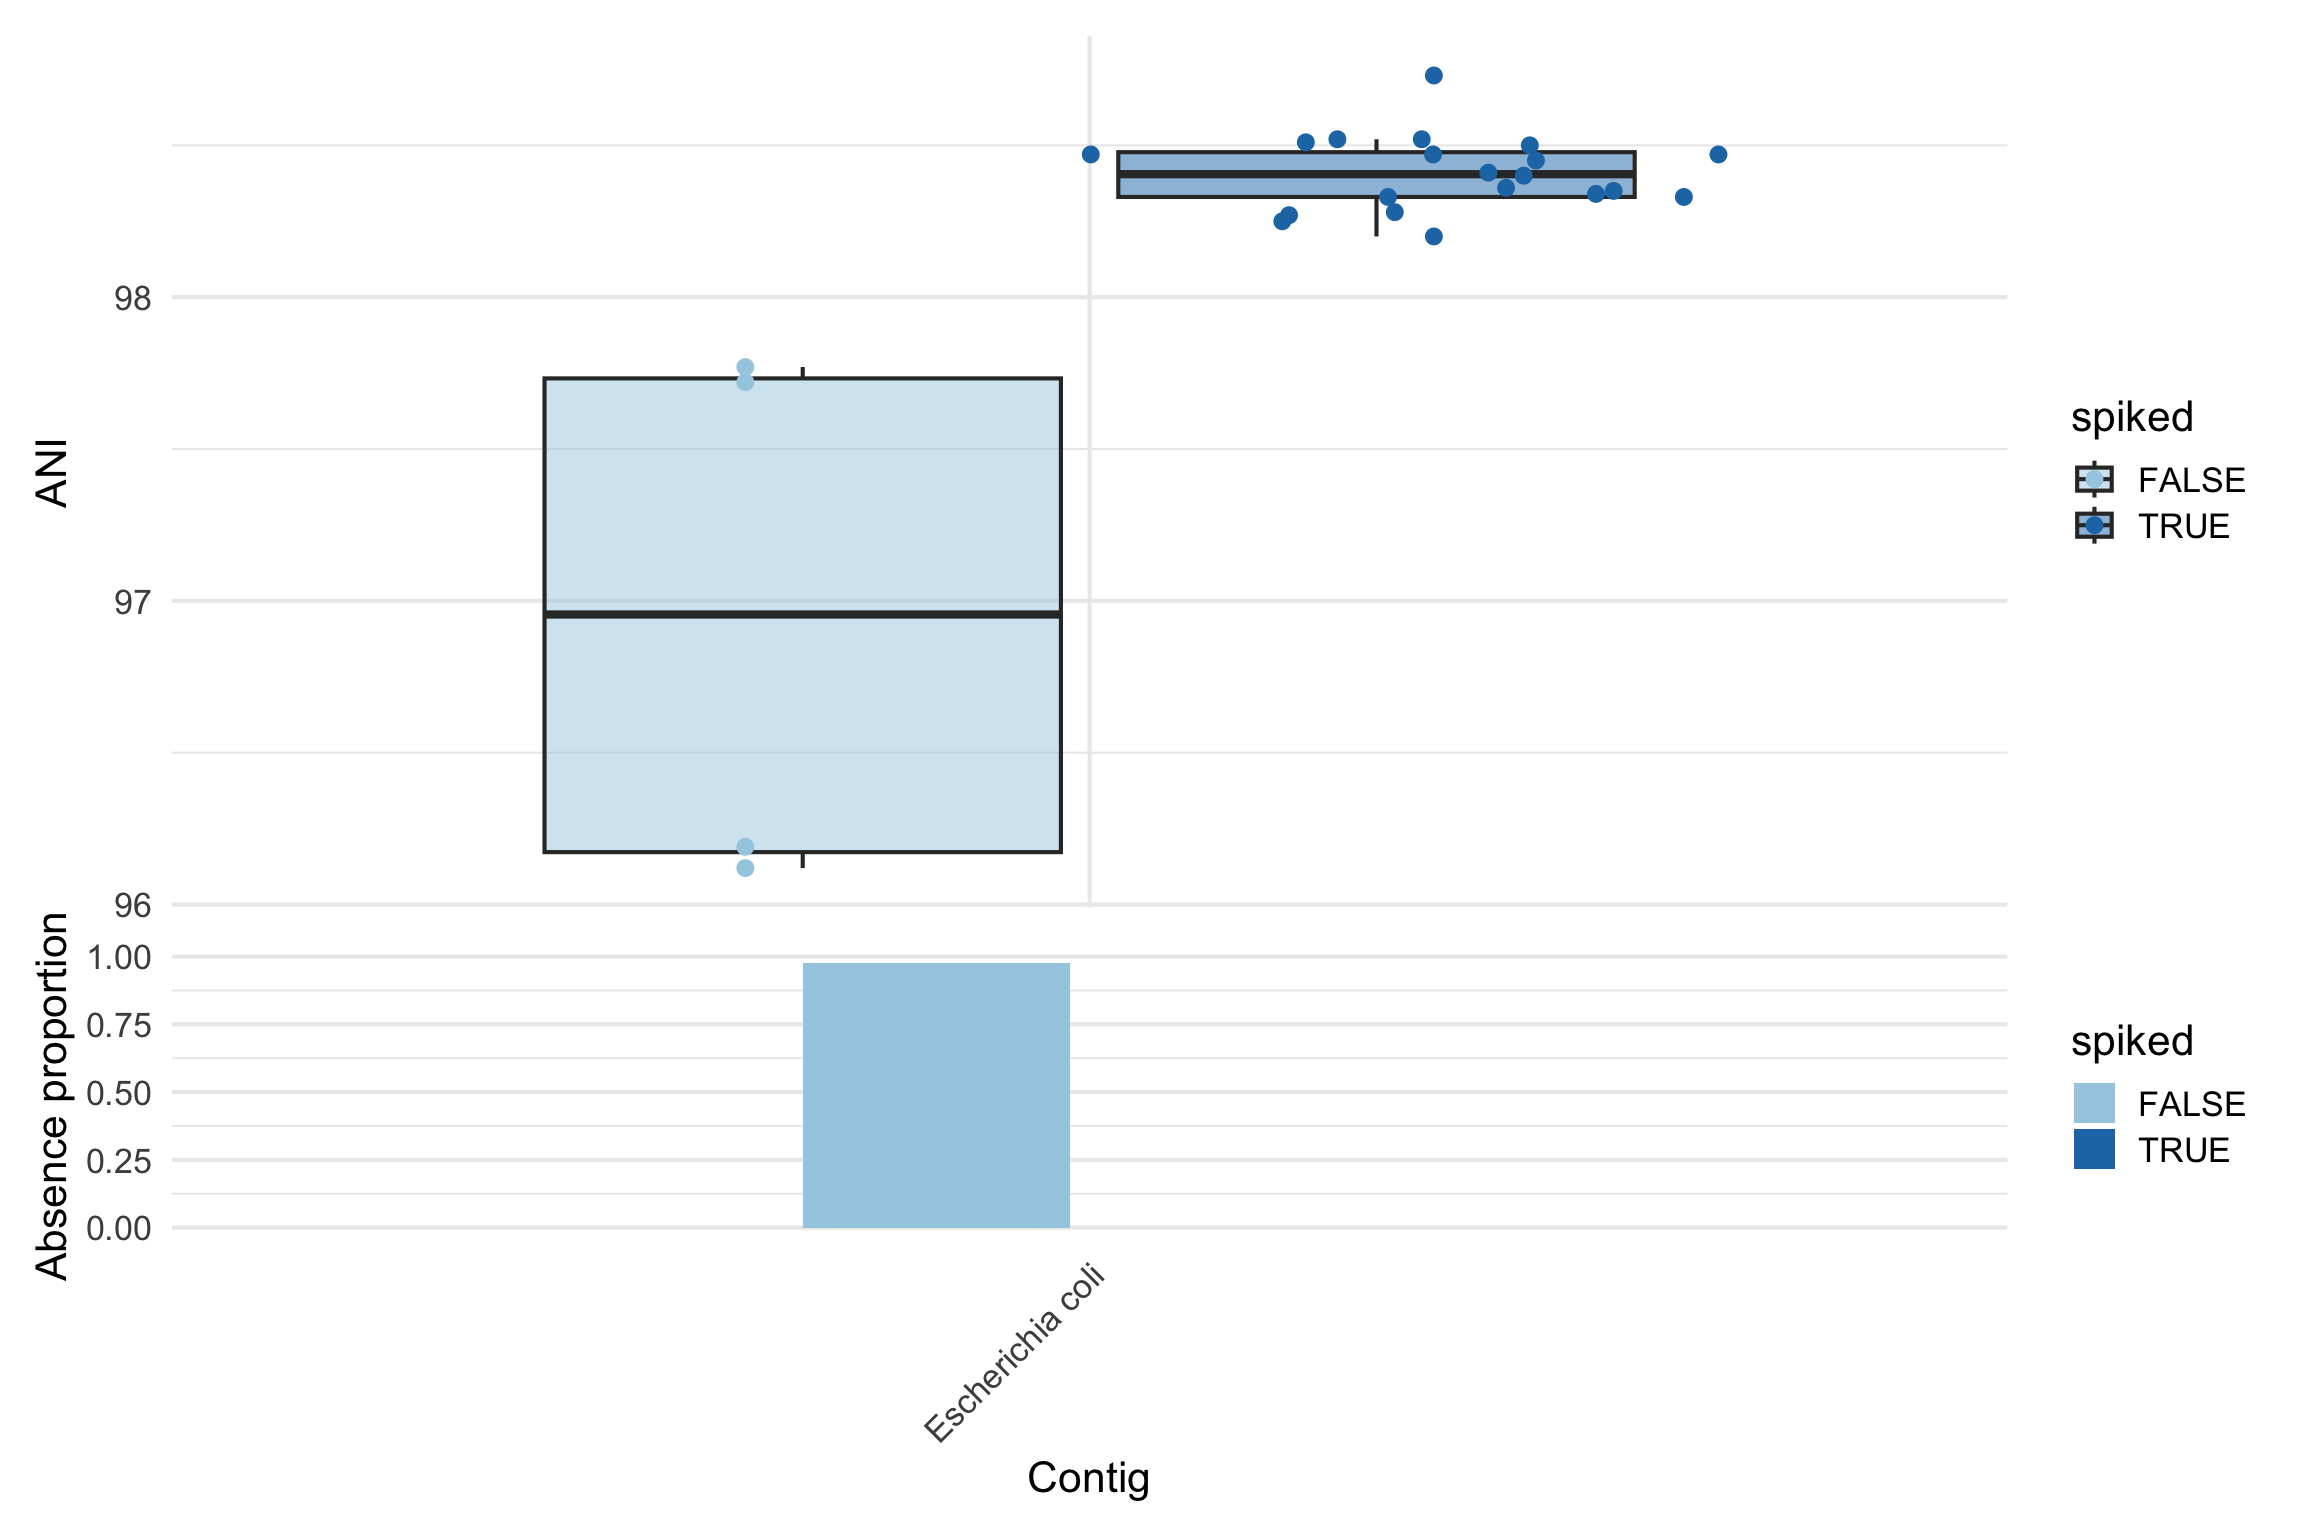
Using the eBayes prior fixed the false positives